return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Cyclobutylamine)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-211.917768
Energy at 298.15K-211.929046
HF Energy-211.142214
Nuclear repulsion energy188.199004
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3560 3344 0.42      
2 A' 3238 3041 32.77      
3 A' 3202 3007 26.63      
4 A' 3160 2968 38.28      
5 A' 3137 2947 18.90      
6 A' 3075 2888 79.07      
7 A' 1690 1587 32.90      
8 A' 1549 1455 4.51      
9 A' 1521 1428 2.94      
10 A' 1416 1330 29.04      
11 A' 1324 1243 8.76      
12 A' 1265 1188 0.69      
13 A' 1190 1118 6.79      
14 A' 1114 1046 8.16      
15 A' 988 928 5.15      
16 A' 920 864 15.30      
17 A' 899 844 47.89      
18 A' 840 789 85.92      
19 A' 679 637 2.94      
20 A' 415 389 4.50      
21 A' 202 189 1.61      
22 A" 3669 3446 2.56      
23 A" 3205 3010 12.19      
24 A" 3133 2943 68.51      
25 A" 1515 1422 2.02      
26 A" 1373 1289 0.01      
27 A" 1304 1224 1.03      
28 A" 1285 1207 0.28      
29 A" 1243 1168 0.81      
30 A" 1183 1111 0.57      
31 A" 1048 984 0.07      
32 A" 970 911 0.67      
33 A" 947 889 2.36      
34 A" 794 746 0.58      
35 A" 398 374 11.64      
36 A" 286 268 30.92      

Unscaled Zero Point Vibrational Energy (zpe) 28865.6 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 27110.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.27680 0.16110 0.13539

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.466 0.757 0.000
H2 -1.833 1.233 0.817
H3 -1.833 1.233 -0.817
C4 0.535 -0.183 -1.071
H5 -0.076 -0.321 -1.962
H6 1.549 0.087 -1.365
C7 0.535 -0.183 1.071
H8 -0.076 -0.321 1.962
H9 1.549 0.087 1.365
C10 -0.005 0.787 0.000
H11 0.417 1.798 0.000
C12 0.535 -1.296 0.000
H13 1.366 -1.999 0.000
H14 -0.404 -1.843 0.000

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 C7 H8 H9 C10 H11 C12 H13 H14
N11.01471.01472.45642.63503.37692.45642.63503.37691.46082.15072.86703.95192.8085
H21.01471.63473.34363.63704.18552.77112.61003.61312.05162.45933.56014.62083.4889
H31.01471.63472.77112.61003.61313.34363.63704.18552.05162.45933.56014.62083.4889
C42.45643.34362.77111.08971.09022.14173.09732.65281.54232.25481.54452.26602.1869
H52.63503.63702.61001.08971.77903.09733.92483.72612.25432.92932.27522.95792.5050
H63.37694.18553.61311.09021.77902.65283.72612.73102.18402.46432.19242.50003.0666
C72.45642.77113.34362.14173.09732.65281.08971.09021.54232.25481.54452.26602.1869
H82.63502.61003.63703.09733.92483.72611.08971.77902.25432.92932.27522.95792.5050
H93.37693.61314.18552.65283.72612.73101.09021.77902.18402.46432.19242.50003.0666
C101.46082.05162.05161.54232.25432.18401.54232.25432.18401.09552.15163.10482.6593
H112.15072.45932.45932.25482.92932.46432.25482.92932.46431.09553.09603.91353.7317
C122.86703.56013.56011.54452.27522.19241.54452.27522.19242.15163.09601.08861.0864
H133.95194.62084.62082.26602.95792.50002.26602.95792.50003.10483.91351.08861.7771
H142.80853.48893.48892.18692.50503.06662.18692.50503.06662.65933.73171.08641.7771

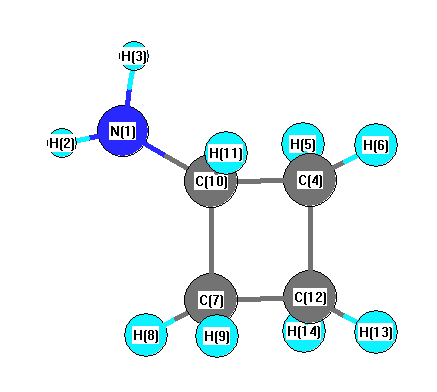
picture of Cyclobutylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C10 C4 109.730 N1 C10 C7 109.730
N1 C10 H11 113.792 H2 N1 H3 107.316
H2 N1 C10 110.651 H3 N1 C10 110.651
C4 C10 C7 87.942 C4 C10 H11 116.448
C4 C12 C7 87.787 C4 C12 H13 117.727
C4 C12 H14 111.255 H5 C4 H6 109.392
H5 C4 C10 116.798 H5 C4 C12 118.448
H6 C4 C10 110.953 H6 C4 C12 111.474
C7 C10 H11 116.448 C7 C12 H13 117.727
C7 C12 H14 111.255 H8 C7 H9 109.392
H8 C7 C10 116.798 H8 C7 C12 118.448
H9 C7 C10 110.953 H9 C7 C12 111.474
C10 C4 C12 88.378 C10 C7 C12 88.378
H13 C12 H14 109.584
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability