Jump to
S1C2
Energy calculated at LSDA/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -417.051939 |
| Energy at 298.15K | -417.055980 |
| HF Energy | -417.051939 |
| Nuclear repulsion energy | 61.135408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at LSDA/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3746 |
3689 |
67.41 |
|
|
|
| 2 |
A' |
2238 |
2205 |
134.98 |
|
|
|
| 3 |
A' |
1090 |
1073 |
14.46 |
|
|
|
| 4 |
A' |
1019 |
1004 |
40.01 |
|
|
|
| 5 |
A' |
871 |
858 |
10.49 |
|
|
|
| 6 |
A' |
782 |
771 |
188.80 |
|
|
|
| 7 |
A" |
2255 |
2221 |
146.70 |
|
|
|
| 8 |
A" |
870 |
857 |
14.79 |
|
|
|
| 9 |
A" |
398 |
392 |
127.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6634.2 cm
-1
Scaled (by 0.985) Zero Point Vibrational Energy (zpe) 6534.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at LSDA/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
-0.109 |
-0.572 |
0.000 |
| O2 |
-0.109 |
1.094 |
0.000 |
| H3 |
0.769 |
1.518 |
0.000 |
| H4 |
0.868 |
-0.846 |
1.032 |
| H5 |
0.868 |
-0.846 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6652 | 2.2666 | 1.4474 | 1.4474 |
O2 | 1.6652 | | 0.9748 | 2.4046 | 2.4046 | H3 | 2.2666 | 0.9748 | | 2.5819 | 2.5819 | H4 | 1.4474 | 2.4046 | 2.5819 | | 2.0649 | H5 | 1.4474 | 2.4046 | 2.5819 | 2.0649 | |
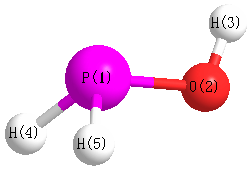 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
115.829 |
|
O2 |
P1 |
H4 |
100.934 |
| O2 |
P1 |
H5 |
100.934 |
|
H4 |
P1 |
H5 |
91.013 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.248 |
|
|
|
| 2 |
O |
-0.635 |
|
|
|
| 3 |
H |
0.379 |
|
|
|
| 4 |
H |
0.004 |
|
|
|
| 5 |
H |
0.004 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.455 |
-0.044 |
0.000 |
2.456 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.666 |
1.904 |
0.000 |
| y |
1.904 |
-20.007 |
0.000 |
| z |
0.000 |
0.000 |
-20.546 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.389 |
1.904 |
0.000 |
| y |
1.904 |
0.598 |
0.000 |
| z |
0.000 |
0.000 |
-0.209 |
|
| Polar |
|---|
| 3z2-r2 | -0.419 |
|---|
| x2-y2 | -0.658 |
|---|
| xy | 1.904 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.160 |
-0.321 |
0.000 |
| y |
-0.321 |
4.412 |
0.000 |
| z |
0.000 |
0.000 |
4.126 |
<r2> (average value of r
2) Å
2
| <r2> |
35.451 |
| (<r2>)1/2 |
5.954 |
Jump to
S1C1
Energy calculated at LSDA/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -417.053459 |
| Energy at 298.15K | -417.057348 |
| HF Energy | -417.053459 |
| Nuclear repulsion energy | 61.039783 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at LSDA/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3779 |
3722 |
123.16 |
|
|
|
| 2 |
A' |
2298 |
2264 |
96.79 |
|
|
|
| 3 |
A' |
1098 |
1081 |
27.65 |
|
|
|
| 4 |
A' |
1077 |
1061 |
84.33 |
|
|
|
| 5 |
A' |
851 |
838 |
41.66 |
|
|
|
| 6 |
A' |
782 |
770 |
112.38 |
|
|
|
| 7 |
A" |
2308 |
2273 |
104.02 |
|
|
|
| 8 |
A" |
890 |
877 |
0.88 |
|
|
|
| 9 |
A" |
257 |
253 |
122.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6669.1 cm
-1
Scaled (by 0.985) Zero Point Vibrational Energy (zpe) 6569.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at LSDA/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
0.040 |
-0.578 |
0.000 |
| O2 |
0.040 |
1.099 |
0.000 |
| H3 |
0.959 |
1.422 |
0.000 |
| H4 |
-0.944 |
-0.772 |
1.032 |
| H5 |
-0.944 |
-0.772 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6774 | 2.2010 | 1.4392 | 1.4392 |
O2 | 1.6774 | | 0.9738 | 2.3526 | 2.3526 | H3 | 2.2010 | 0.9738 | | 3.0821 | 3.0821 | H4 | 1.4392 | 2.3526 | 3.0821 | | 2.0639 | H5 | 1.4392 | 2.3526 | 3.0821 | 2.0639 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
109.348 |
|
O2 |
P1 |
H4 |
97.736 |
| O2 |
P1 |
H5 |
97.736 |
|
H4 |
P1 |
H5 |
91.623 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.224 |
|
|
|
| 2 |
O |
-0.662 |
|
|
|
| 3 |
H |
0.389 |
|
|
|
| 4 |
H |
0.024 |
|
|
|
| 5 |
H |
0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.742 |
-0.237 |
0.000 |
0.779 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.552 |
4.346 |
0.000 |
| y |
4.346 |
-20.856 |
0.000 |
| z |
0.000 |
0.000 |
-20.419 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.086 |
4.346 |
0.000 |
| y |
4.346 |
-0.871 |
0.000 |
| z |
0.000 |
0.000 |
-0.215 |
|
| Polar |
|---|
| 3z2-r2 | -0.429 |
|---|
| x2-y2 | 1.304 |
|---|
| xy | 4.346 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.266 |
0.414 |
0.000 |
| y |
0.414 |
4.470 |
0.000 |
| z |
0.000 |
0.000 |
4.134 |
<r2> (average value of r
2) Å
2
| <r2> |
35.427 |
| (<r2>)1/2 |
5.952 |
