Jump to
S1C2
Energy calculated at B97D3/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -418.325044 |
| Energy at 298.15K | -418.329085 |
| HF Energy | -418.325044 |
| Nuclear repulsion energy | 60.592750 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3745 |
3681 |
45.30 |
|
|
|
| 2 |
A' |
2255 |
2216 |
168.28 |
|
|
|
| 3 |
A' |
1122 |
1103 |
20.96 |
|
|
|
| 4 |
A' |
1066 |
1048 |
35.06 |
|
|
|
| 5 |
A' |
886 |
870 |
40.97 |
|
|
|
| 6 |
A' |
733 |
720 |
151.85 |
|
|
|
| 7 |
A" |
2265 |
2226 |
188.23 |
|
|
|
| 8 |
A" |
876 |
861 |
14.01 |
|
|
|
| 9 |
A" |
400 |
393 |
117.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6673.8 cm
-1
Scaled (by 0.9828) Zero Point Vibrational Energy (zpe) 6559.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
-0.109 |
-0.580 |
0.000 |
| O2 |
-0.109 |
1.111 |
0.000 |
| H3 |
0.775 |
1.510 |
0.000 |
| H4 |
0.862 |
-0.849 |
1.032 |
| H5 |
0.862 |
-0.849 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6908 | 2.2694 | 1.4424 | 1.4424 |
O2 | 1.6908 | | 0.9700 | 2.4185 | 2.4185 | H3 | 2.2694 | 0.9700 | | 2.5766 | 2.5766 | H4 | 1.4424 | 2.4185 | 2.5766 | | 2.0642 | H5 | 1.4424 | 2.4185 | 2.5766 | 2.0642 | |
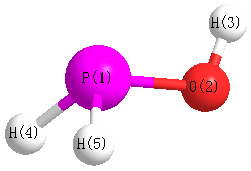 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
116.259 |
|
O2 |
P1 |
H4 |
101.360 |
| O2 |
P1 |
H5 |
101.360 |
|
H4 |
P1 |
H5 |
91.860 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.338 |
|
|
|
| 2 |
O |
-0.635 |
|
|
|
| 3 |
H |
0.358 |
|
|
|
| 4 |
H |
-0.031 |
|
|
|
| 5 |
H |
-0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.309 |
-0.216 |
0.000 |
2.320 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.527 |
1.882 |
0.000 |
| y |
1.882 |
-20.084 |
0.000 |
| z |
0.000 |
0.000 |
-20.422 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.274 |
1.882 |
0.000 |
| y |
1.882 |
0.390 |
0.000 |
| z |
0.000 |
0.000 |
-0.117 |
|
| Polar |
|---|
| 3z2-r2 | -0.234 |
|---|
| x2-y2 | -0.443 |
|---|
| xy | 1.882 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.096 |
-0.313 |
0.000 |
| y |
-0.313 |
4.451 |
0.000 |
| z |
0.000 |
0.000 |
4.098 |
<r2> (average value of r
2) Å
2
| <r2> |
35.837 |
| (<r2>)1/2 |
5.986 |
Jump to
S1C1
Energy calculated at B97D3/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -418.326246 |
| Energy at 298.15K | -418.330116 |
| HF Energy | -418.326246 |
| Nuclear repulsion energy | 60.486230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3779 |
3714 |
88.34 |
|
|
|
| 2 |
A' |
2310 |
2270 |
127.59 |
|
|
|
| 3 |
A' |
1128 |
1108 |
23.10 |
|
|
|
| 4 |
A' |
1112 |
1093 |
102.03 |
|
|
|
| 5 |
A' |
874 |
859 |
19.62 |
|
|
|
| 6 |
A' |
726 |
714 |
115.21 |
|
|
|
| 7 |
A" |
2314 |
2274 |
140.70 |
|
|
|
| 8 |
A" |
897 |
881 |
0.97 |
|
|
|
| 9 |
A" |
246 |
241 |
112.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6692.8 cm
-1
Scaled (by 0.9828) Zero Point Vibrational Energy (zpe) 6577.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
0.040 |
-0.587 |
0.000 |
| O2 |
0.040 |
1.117 |
0.000 |
| H3 |
0.958 |
1.425 |
0.000 |
| H4 |
-0.937 |
-0.778 |
1.032 |
| H5 |
-0.937 |
-0.778 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.7032 | 2.2114 | 1.4340 | 1.4340 |
O2 | 1.7032 | | 0.9688 | 2.3686 | 2.3686 | H3 | 2.2114 | 0.9688 | | 3.0841 | 3.0841 | H4 | 1.4340 | 2.3686 | 3.0841 | | 2.0647 | H5 | 1.4340 | 2.3686 | 3.0841 | 2.0647 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
111.652 |
|
O2 |
P1 |
H4 |
96.928 |
| O2 |
P1 |
H5 |
96.928 |
|
H4 |
P1 |
H5 |
92.359 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.316 |
|
|
|
| 2 |
O |
-0.659 |
|
|
|
| 3 |
H |
0.366 |
|
|
|
| 4 |
H |
-0.012 |
|
|
|
| 5 |
H |
-0.012 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.760 |
-0.381 |
0.000 |
0.850 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.510 |
4.138 |
0.000 |
| y |
4.138 |
-20.817 |
0.000 |
| z |
0.000 |
0.000 |
-20.295 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.046 |
4.138 |
0.000 |
| y |
4.138 |
-0.915 |
0.000 |
| z |
0.000 |
0.000 |
-0.132 |
|
| Polar |
|---|
| 3z2-r2 | -0.264 |
|---|
| x2-y2 | 1.307 |
|---|
| xy | 4.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.197 |
0.412 |
0.000 |
| y |
0.412 |
4.541 |
0.000 |
| z |
0.000 |
0.000 |
4.104 |
<r2> (average value of r
2) Å
2
| <r2> |
35.842 |
| (<r2>)1/2 |
5.987 |
