Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -131.031806 |
| Energy at 298.15K | -131.031398 |
| HF Energy | -130.682787 |
| Nuclear repulsion energy | 47.382450 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3402 |
3195 |
15.29 |
|
|
|
| 2 |
A' |
2282 |
2144 |
1.17 |
|
|
|
| 3 |
A' |
1079 |
1013 |
31.94 |
|
|
|
| 4 |
A' |
854 |
802 |
5.96 |
|
|
|
| 5 |
A' |
442 |
415 |
4.54 |
|
|
|
| 6 |
A" |
507 |
476 |
6.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4283.1 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4022.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.268 |
-1.231 |
0.000 |
| C2 |
0.000 |
0.119 |
0.000 |
| N3 |
0.183 |
1.256 |
0.000 |
| H4 |
0.329 |
-2.122 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3771 | 2.5282 | 1.0724 |
C2 | 1.3771 | | 1.1516 | 2.2653 | N3 | 2.5282 | 1.1516 | | 3.3814 | H4 | 1.0724 | 2.2653 | 3.3814 | |
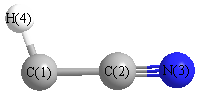 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
177.909 |
|
C2 |
C1 |
H4 |
134.917 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -131.025830 |
| Energy at 298.15K | |
| HF Energy | -130.682519 |
| Nuclear repulsion energy | 47.671257 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3519 |
3305 |
78.78 |
|
|
|
| 2 |
Σ |
1705 |
1601 |
37.20 |
|
|
|
| 3 |
Σ |
1207 |
1133 |
46.07 |
|
|
|
| 4 |
Π |
493 |
463 |
0.41 |
|
|
|
| 4 |
Π |
493 |
463 |
0.41 |
|
|
|
| 5 |
Π |
577i |
542i |
69.76 |
|
|
|
| 5 |
Π |
577i |
542i |
69.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3131.3 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 2940.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.212 |
| C2 |
0.000 |
0.000 |
0.099 |
| N3 |
0.000 |
0.000 |
1.279 |
| H4 |
0.000 |
0.000 |
-2.274 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3107 | 2.4902 | 1.0628 |
C2 | 1.3107 | | 1.1794 | 2.3735 | N3 | 2.4902 | 1.1794 | | 3.5530 | H4 | 1.0628 | 2.3735 | 3.5530 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31+G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.178 |
|
|
|
| 2 |
C |
-0.037 |
|
|
|
| 3 |
N |
-0.424 |
|
|
|
| 4 |
H |
0.282 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.800 |
3.800 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.716 |
0.000 |
0.000 |
| y |
0.000 |
-17.716 |
0.000 |
| z |
0.000 |
0.000 |
-16.150 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.783 |
0.000 |
0.000 |
| y |
0.000 |
-0.783 |
0.000 |
| z |
0.000 |
0.000 |
1.566 |
|
| Polar |
|---|
| 3z2-r2 | 3.132 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.147 |
0.000 |
0.000 |
| y |
0.000 |
2.147 |
0.000 |
| z |
0.000 |
0.000 |
6.963 |
<r2> (average value of r
2) Å
2
| <r2> |
36.222 |
| (<r2>)1/2 |
6.018 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -131.025979 |
| Energy at 298.15K | -131.025377 |
| HF Energy | -130.621360 |
| Nuclear repulsion energy | 46.472851 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3145 |
2954 |
13.36 |
|
|
|
| 2 |
A' |
2254 |
2117 |
8.17 |
|
|
|
| 3 |
A' |
1069 |
1004 |
78.91 |
|
|
|
| 4 |
A' |
989 |
929 |
2.78 |
|
|
|
| 5 |
A' |
385 |
361 |
18.24 |
|
|
|
| 6 |
A" |
325 |
306 |
8.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4083.0 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 3834.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.233 |
-1.286 |
0.000 |
| C2 |
0.000 |
0.100 |
0.000 |
| N3 |
-0.387 |
1.224 |
0.000 |
| H4 |
1.316 |
-1.455 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4055 | 2.5852 | 1.0961 |
C2 | 1.4055 | | 1.1886 | 2.0369 | N3 | 2.5852 | 1.1886 | | 3.1742 | H4 | 1.0961 | 2.0369 | 3.1742 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -131.025979 |
| Energy at 298.15K | -131.025377 |
| HF Energy | -130.621360 |
| Nuclear repulsion energy | 46.472851 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
