return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (1H-Pyrazole)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-225.575542
Energy at 298.15K-225.581444
HF Energy-224.807784
Nuclear repulsion energy 
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3730 3503 101.21      
2 A' 3361 3156 0.11      
3 A' 3344 3140 1.11      
4 A' 3335 3132 0.99      
5 A' 1571 1476 5.54      
6 A' 1520 1427 7.49      
7 A' 1481 1391 10.99      
8 A' 1395 1310 4.10      
9 A' 1305 1226 2.09      
10 A' 1198 1125 1.17      
11 A' 1174 1103 14.43      
12 A' 1082 1016 13.21      
13 A' 1075 1010 30.11      
14 A' 938 881 2.19      
15 A' 917 861 6.74      
16 A" 827 777 27.89      
17 A" 786 739 24.17      
18 A" 718 674 93.06      
19 A" 680 639 30.05      
20 A" 600 564 0.87      
21 A" 501 471 41.73      

Unscaled Zero Point Vibrational Energy (zpe) 15769.0 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 14810.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.32019 0.31330 0.15835

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 2.104 0.732 0.000
C2 1.116 0.308 0.000
H3 1.262 -1.894 0.000
C4 0.661 -1.001 0.000
H5 -1.477 -1.684 0.000
C6 -0.741 -0.899 0.000
N7 -1.151 0.386 0.000
H8 -0.048 2.093 0.000
N9 0.000 1.087 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 C6 N7 H8 N9
H11.07602.75752.25544.32033.27953.27372.54632.1340
C21.07602.20631.38573.26952.21412.26792.13081.3607
H32.75752.20631.07552.74732.23633.31954.19633.2367
C42.25541.38571.07552.24501.40602.28243.17472.1906
H54.32033.26952.74732.24501.07642.09534.03843.1400
C63.27952.21412.23631.40601.07641.34873.07122.1195
N73.27372.26793.31952.28242.09531.34872.03261.3477
H82.54632.13084.19633.17474.03843.07122.03261.0072
N92.13401.36073.23672.19063.14002.11951.34771.0072

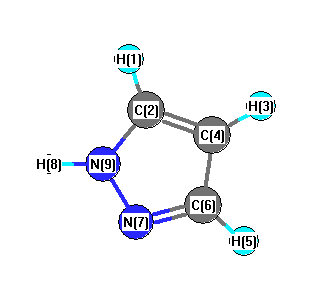
picture of 1H-Pyrazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 132.354 H1 C2 N9 121.843
C2 C4 H3 126.926 C2 C4 C6 104.949
C2 N9 N7 113.723 C2 N9 H8 127.652
H3 C4 C6 128.125 C4 C2 N9 105.803
C4 C6 H5 128.995 C4 C6 N7 111.887
H5 C6 N7 119.118 C6 N7 N9 103.638
N7 N9 H8 118.625
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability