return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (1H-Imidazole)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-225.591825
Energy at 298.15K-225.597704
HF Energy-224.829895
Nuclear repulsion energy162.897237
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3726 3500 77.38      
2 A' 3363 3158 0.79      
3 A' 3344 3141 0.85      
4 A' 3338 3135 2.63      
5 A' 1563 1468 6.48      
6 A' 1528 1435 26.29      
7 A' 1489 1399 19.22      
8 A' 1398 1313 8.59      
9 A' 1300 1221 1.11      
10 A' 1194 1121 2.76      
11 A' 1171 1100 3.69      
12 A' 1123 1055 26.79      
13 A' 1099 1032 29.61      
14 A' 940 883 2.50      
15 A' 901 846 4.78      
16 A" 814 765 26.96      
17 A" 768 721 61.50      
18 A" 693 651 25.12      
19 A" 670 629 0.01      
20 A" 620 582 33.95      
21 A" 487 457 74.86      

Unscaled Zero Point Vibrational Energy (zpe) 15763.6 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 14805.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.32505 0.31134 0.15902

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 1.105 0.000
C2 -1.093 0.285 0.000
C3 1.120 0.306 0.000
N4 -0.740 -0.993 0.000
C5 0.636 -0.985 0.000
H6 -0.012 2.112 0.000
H7 -2.101 0.662 0.000
H8 2.116 0.711 0.000
H9 1.201 -1.901 0.000

Atom - Atom Distances (Å)
  N1 C2 C3 N4 C5 H6 H7 H8 H9
N11.36631.37542.22462.18471.00742.14692.15203.2369
C21.36632.21281.32582.14572.12291.07623.23683.1690
C31.37542.21282.26871.37882.13163.24021.07512.2084
N42.22461.32582.26871.37653.18942.14293.32562.1434
C52.18472.14571.37881.37653.16463.19482.25061.0760
H61.00742.12292.13163.18943.16462.54242.54794.1926
H72.14691.07623.24022.14293.19482.54244.21684.1804
H82.15203.23681.07513.32562.25062.54794.21682.7671
H93.23693.16902.20842.14341.07604.19264.18042.7671

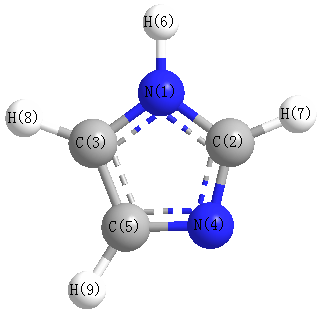
picture of 1H-Imidazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 N4 111.442 N1 C2 H7 122.596
N1 C3 C5 104.977 N1 C3 H8 122.376
C2 N1 C3 107.624 C2 N1 H6 126.173
C2 N4 C5 105.105 C3 N1 H6 126.203
C3 C5 N4 110.852 C3 C5 H9 127.791
N4 C2 H7 125.962 N4 C5 H9 121.357
C5 C3 H8 132.647
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability