return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5SC2H5 (Diethyl sulfide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-555.575736
Energy at 298.15K-555.586333
HF Energy-554.822498
Nuclear repulsion energy234.184805
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3220 3024 30.37      
2 A1 3125 2935 0.82      
3 A1 3119 2929 60.79      
4 A1 1548 1454 4.61      
5 A1 1534 1441 0.38      
6 A1 1462 1373 1.55      
7 A1 1372 1288 13.26      
8 A1 1135 1066 0.79      
9 A1 1036 973 2.58      
10 A1 734 690 0.00      
11 A1 343 322 0.81      
12 A1 147 138 0.17      
13 A2 3229 3033 0.00      
14 A2 3183 2990 0.00      
15 A2 1536 1443 0.00      
16 A2 1294 1215 0.00      
17 A2 1069 1004 0.00      
18 A2 807 758 0.00      
19 A2 259 244 0.00      
20 A2 48 45 0.00      
21 B1 3229 3032 41.91      
22 B1 3177 2984 18.62      
23 B1 1536 1443 18.00      
24 B1 1294 1216 0.15      
25 B1 1092 1026 0.22      
26 B1 823 773 5.53      
27 B1 256 240 0.15      
28 B1 52 49 0.78      
29 B2 3220 3024 10.89      
30 B2 3125 2935 48.74      
31 B2 3121 2932 0.86      
32 B2 1547 1453 1.14      
33 B2 1528 1435 9.84      
34 B2 1459 1370 5.53      
35 B2 1340 1258 41.27      
36 B2 1087 1021 1.95      
37 B2 1032 969 3.31      
38 B2 737 692 0.57      
39 B2 347 326 0.98      

Unscaled Zero Point Vibrational Energy (zpe) 30100.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 28270.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.48032 0.05793 0.05377

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.565
C2 0.000 1.376 -0.614
C3 0.000 -1.376 -0.614
C4 0.000 2.706 0.124
C5 0.000 -2.706 0.124
H6 0.883 1.301 -1.250
H7 -0.883 1.301 -1.250
H8 0.883 -1.301 -1.250
H9 -0.883 -1.301 -1.250
H10 0.000 3.529 -0.589
H11 0.000 -3.529 -0.589
H12 -0.882 2.802 0.754
H13 0.882 2.802 0.754
H14 0.882 -2.802 0.754
H15 -0.882 -2.802 0.754

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.81211.81212.74142.74142.40122.40122.40122.40123.71263.71262.94402.94402.94402.9440
C21.81212.75161.52134.14781.09071.09072.88932.88932.15314.90462.16472.16474.48424.4842
C31.81212.75164.14781.52132.88932.88931.09071.09074.90462.15314.48424.48422.16472.1647
C42.74141.52134.14785.41162.15452.15454.32674.32671.08906.27521.08831.08835.61385.6138
C52.74144.14781.52135.41164.32674.32672.15452.15456.27521.08905.61385.61381.08831.0883
H62.40121.09072.88932.15454.32671.76662.60163.14472.48614.95393.06392.50414.56644.8958
H72.40121.09072.88932.15454.32671.76663.14472.60162.48614.95392.50413.06394.89584.5664
H82.40122.88931.09074.32672.15452.60163.14471.76664.95392.48614.89584.56642.50413.0639
H92.40122.88931.09074.32672.15453.14472.60161.76664.95392.48614.56644.89583.06392.5041
H103.71262.15314.90461.08906.27522.48612.48614.95394.95397.05751.76351.76356.53196.5319
H113.71264.90462.15316.27521.08904.95394.95392.48612.48617.05756.53196.53191.76351.7635
H122.94402.16474.48421.08835.61383.06392.50414.89584.56641.76356.53191.76435.87595.6047
H132.94402.16474.48421.08835.61382.50413.06394.56644.89581.76356.53191.76435.60475.8759
H142.94404.48422.16475.61381.08834.56644.89582.50413.06396.53191.76355.87595.60471.7643
H152.94404.48422.16475.61381.08834.89584.56643.06392.50416.53191.76355.60475.87591.7643

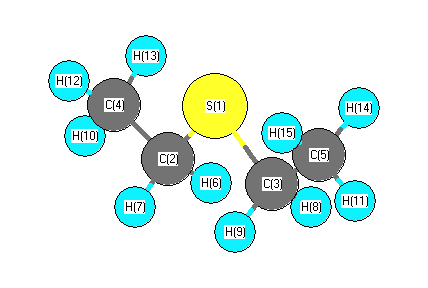
picture of Diethyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C4 110.354 S1 C2 H6 109.086
S1 C2 H7 109.086 S1 C3 C5 110.354
S1 C3 H8 109.086 S1 C3 H9 109.086
C2 S1 C3 98.795 C2 C4 H10 110.045
C2 C4 H12 111.015 C2 C4 H13 111.015
C3 C5 H11 110.045 C3 C5 H14 111.015
C3 C5 H15 111.015 C4 C2 H6 110.058
C4 C2 H7 110.058 C5 C3 H8 110.058
C5 C3 H9 110.058 H6 C2 H7 108.156
H8 C3 H9 108.156 H10 C4 H12 108.181
H10 C4 H13 108.181 H11 C5 H14 108.181
H11 C5 H15 108.181 H12 C4 H13 108.306
H14 C5 H15 108.306
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability