return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-1272.490231
Energy at 298.15K-1272.496540
Nuclear repulsion energy365.348263
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3250 3052 5.44      
2 A 3239 3042 0.06      
3 A 3129 2939 3.46      
4 A 1518 1425 7.87      
5 A 1493 1403 9.42      
6 A 1418 1332 0.12      
7 A 1021 958 0.09      
8 A 1018 956 2.60      
9 A 749 704 0.19      
10 A 502 471 0.33      
11 A 273 256 0.38      
12 A 198 186 1.33      
13 A 162 152 1.35      
14 A 64 60 2.11      
15 B 3249 3052 2.28      
16 B 3239 3042 4.63      
17 B 3129 2939 31.95      
18 B 1516 1424 14.06      
19 B 1494 1403 9.30      
20 B 1421 1334 3.71      
21 B 1018 957 2.07      
22 B 1013 951 11.51      
23 B 748 703 0.90      
24 B 500 469 15.07      
25 B 281 264 1.10      
26 B 190 178 0.75      
27 B 95 89 3.51      

Unscaled Zero Point Vibrational Energy (zpe) 17961.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16869.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.15096 0.04611 0.04577

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.075
S2 0.000 1.653 -0.158
S3 0.000 -1.653 -0.158
C4 1.741 1.734 -0.638
C5 -1.741 -1.734 -0.638
H6 1.849 2.600 -1.289
H7 -1.849 -2.600 -1.289
H8 2.379 1.856 0.232
H9 2.026 0.841 -1.189
H10 -2.379 -1.856 0.232
H11 -2.026 -0.841 -1.189

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.06292.06292.99582.99583.97093.97093.13333.15283.13333.1528
S22.06293.30701.80763.83892.36494.77362.41942.41394.25803.3752
S32.06293.30703.83891.80764.77362.36494.25803.37522.41942.4139
C42.99581.80763.83894.91421.08845.66481.08631.08695.53374.5962
C52.99583.83891.80764.91425.66481.08845.53374.59621.08631.0869
H63.97092.36494.77361.08845.66486.38011.77431.77026.32815.1832
H73.97094.77362.36495.66481.08846.38016.32815.18321.77431.7702
H83.13332.41944.25801.08635.53371.77436.32811.78146.03535.3575
H93.15282.41393.37521.08694.59621.77025.18321.78145.35754.3877
H103.13334.25802.41945.53371.08636.32811.77436.03535.35751.7814
H113.15283.37522.41394.59621.08695.18321.77025.35754.38771.7814

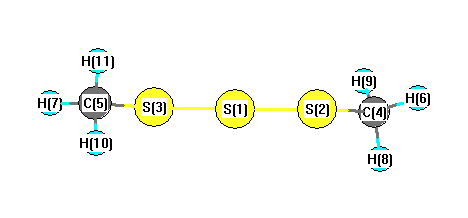
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 101.225 S1 S3 C5 101.225
S2 S1 S3 106.555 S2 C4 H6 106.850
S2 C4 H8 110.981 S2 C4 H9 110.533
S3 C5 H7 106.850 S3 C5 H10 110.981
S3 C5 H11 110.533 H6 C4 H8 109.350
H6 C4 H9 108.929 H7 C5 H10 109.350
H7 C5 H11 108.929 H8 C4 H9 110.116
H10 C5 H11 110.116
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability