return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-429.589497
Energy at 298.15K-429.593836
Nuclear repulsion energy346.268215
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3293 3093 1.95      
2 A1 3280 3080 6.16      
3 A1 1680 1578 25.04      
4 A1 1565 1470 174.14      
5 A1 1471 1382 2.25      
6 A1 1311 1231 125.11      
7 A1 1197 1124 1.41      
8 A1 1060 996 8.02      
9 A1 775 728 35.80      
10 A1 579 544 4.40      
11 A1 283 266 0.30      
12 A2 891 837 0.00      
13 A2 813 764 0.00      
14 A2 535 503 0.00      
15 A2 310 291 0.00      
16 A2 172 162 0.00      
17 B1 888 834 4.33      
18 B1 756 710 102.55      
19 B1 440 413 2.67      
20 B1 286 269 0.01      
21 B2 3288 3088 1.29      
22 B2 3269 3070 1.13      
23 B2 1675 1573 10.26      
24 B2 1509 1418 10.89      
25 B2 1302 1223 2.55      
26 B2 1226 1151 35.19      
27 B2 1132 1063 30.83      
28 B2 861 809 16.99      
29 B2 548 515 3.63      
30 B2 438 412 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 18416.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 17297.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.10813 0.07369 0.04382

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.695 -0.533
C2 0.000 -0.695 -0.533
C3 0.000 -1.399 0.663
C4 0.000 -0.699 1.869
C5 0.000 0.699 1.869
C6 0.000 1.399 0.663
F7 0.000 1.356 -1.713
F8 0.000 -1.356 -1.713
H9 0.000 -2.479 0.629
H10 0.000 -1.244 2.801
H11 0.000 1.244 2.801
H12 0.000 2.479 0.629

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.39052.41182.77722.40211.38771.35232.36633.38033.85733.37932.1289
C21.39051.38772.40212.77722.41182.36631.35232.12893.37933.85733.3803
C32.41181.38771.39492.41972.79833.63802.37641.08022.14423.39973.8781
C42.77722.40211.39491.39702.41974.12943.64202.16941.08012.15443.4106
C52.40212.77722.41971.39701.39493.64204.12943.41062.15441.08012.1694
C61.38772.41182.79832.41971.39492.37643.63803.87813.39972.14421.0802
F71.35232.36633.63804.12943.64202.37642.71174.49355.20954.51602.5978
F82.36631.35232.37643.64204.12943.63802.71172.59784.51605.20954.4935
H93.38032.12891.08022.16943.41063.87814.49352.59782.49874.30994.9576
H103.85733.37932.14421.08012.15443.39975.20954.51602.49872.48744.3099
H113.37933.85733.39972.15441.08012.14424.51605.20954.30992.48742.4987
H122.12893.38033.87813.41062.16941.08022.59784.49354.95764.30992.4987

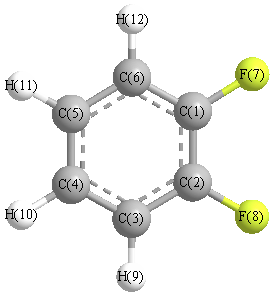
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.479 C1 C2 F8 119.239
C1 C6 C5 119.371 C1 C6 H12 118.704
C2 C1 C6 120.479 C2 C1 F7 119.239
C2 C3 C4 119.371 C2 C3 H9 118.704
C3 C2 F8 120.282 C3 C4 C5 120.150
C3 C4 H10 119.534 C4 C3 H9 121.925
C4 C5 C6 120.150 C4 C5 H11 120.316
C5 C4 H10 120.316 C5 C6 H12 121.925
C6 C1 F7 120.282 C6 C5 H11 119.534
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability