return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-307.447825
Energy at 298.15K 
HF Energy-306.353565
Nuclear repulsion energy312.100219
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3318 3116 10.21      
2 A1 3276 3077 14.77      
3 A1 3260 3062 7.41      
4 A1 1714 1610 0.50      
5 A1 1555 1461 3.75      
6 A1 1535 1442 4.03      
7 A1 1464 1375 7.11      
8 A1 1206 1133 0.02      
9 A1 1143 1073 0.15      
10 A1 1086 1020 0.93      
11 A1 1009 948 4.09      
12 A1 823 773 0.21      
13 A1 549 515 0.08      
14 A2 873 820 0.00      
15 A2 854 802 0.00      
16 A2 790 742 0.00      
17 A2 452 425 0.00      
18 A2 275 258 0.00      
19 A2 500i 469i 0.00      
20 B1 844 793 4.76      
21 B1 726 682 109.47      
22 B1 665 625 44.02      
23 B1 308 289 0.01      
24 B1 139 131 12.67      
25 B2 3289 3089 1.83      
26 B2 3268 3069 26.08      
27 B2 3249 3052 0.16      
28 B2 1664 1563 0.73      
29 B2 1487 1396 2.54      
30 B2 1325 1245 4.77      
31 B2 1274 1196 13.36      
32 B2 1124 1056 2.02      
33 B2 1059 994 1.60      
34 B2 871 818 0.52      
35 B2 655 615 1.34      
36 B2 421 395 3.75      

Unscaled Zero Point Vibrational Energy (zpe) 23524.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 22094.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.16032 0.07325 0.05028

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.446 -0.622
C2 0.000 0.690 -1.844
C3 0.000 -0.690 -1.844
C4 0.000 -1.446 -0.622
C5 0.000 -0.680 2.043
C6 0.000 0.680 2.043
C7 0.000 0.717 0.529
C8 0.000 -0.717 0.529
H9 0.000 2.528 -0.652
H10 0.000 1.224 -2.785
H11 0.000 -1.224 -2.785
H12 0.000 -2.528 -0.652
H13 0.000 -1.446 2.805
H14 0.000 1.446 2.805

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.43722.46112.89263.40952.77361.36322.45021.08252.17393.43643.97484.48413.4269
C21.43721.37982.46114.12183.88762.37362.75892.19161.08172.13273.43225.11604.7100
C32.46111.37981.43723.88764.12182.75892.37363.43222.13271.08172.19164.71005.1160
C42.89262.46111.43722.77363.40952.45021.36323.97483.43642.17391.08253.42694.4841
C53.40954.12183.88762.77361.35912.05961.51464.18965.18994.85873.26801.08002.2574
C62.77363.88764.12183.40951.35911.51462.05963.26804.85875.18994.18962.25741.0800
C71.36322.37362.75892.45022.05961.51461.43312.16263.35263.84043.45303.13892.3895
C82.45022.75892.37361.36321.51462.05961.43313.45303.84043.35262.16262.38953.1389
H91.08252.19163.43223.97484.18963.26802.16263.45302.50044.31665.05685.26663.6218
H102.17391.08172.13273.43645.18994.85873.35263.84042.50042.44854.31666.19445.5939
H113.43642.13271.08172.17394.85875.18993.84043.35264.31662.44852.50045.59396.1944
H123.97483.43222.19161.08253.26804.18963.45302.16265.05684.31662.50043.62185.2666
H134.48415.11604.71003.42691.08002.25743.13892.38955.26666.19445.59393.62182.8911
H143.42694.71005.11604.48412.25741.08002.38953.13893.62185.59396.19445.26662.8911

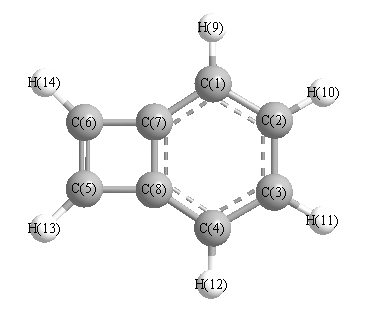
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.756 C1 C2 H10 118.640
C1 C7 C6 149.030 C1 C7 N8 122.368
C2 C1 C7 115.876 C2 C1 H9 120.208
C2 C3 C4 121.756 C2 C3 H11 119.604
C3 C2 H10 119.604 C3 C4 N8 115.876
C3 C4 H12 120.208 C4 C3 H11 118.640
C4 N8 C5 149.030 C4 N8 C7 122.368
C5 C6 C7 91.398 C5 C6 H14 135.172
C5 N8 C7 88.602 C6 C5 N8 91.398
C6 C5 H13 135.172 C6 C7 N8 88.602
C7 C1 H9 123.916 C7 C6 H14 133.430
N8 C4 H12 123.916 N8 C5 H13 133.430
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability