return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NNCH3 ((E)-1,2-Dimethyldiazene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-188.730344
Energy at 298.15K-188.737751
HF Energy-188.080830
Nuclear repulsion energy119.710962
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3231 3034 0.00      
2 Ag 3114 2925 0.00      
3 Ag 1591 1494 0.00      
4 Ag 1517 1425 0.00      
5 Ag 1449 1361 0.00      
6 Ag 1234 1159 0.00      
7 Ag 954 896 0.00      
8 Ag 604 567 0.00      
9 Au 3227 3031 17.93      
10 Au 1522 1430 17.75      
11 Au 1160 1089 3.20      
12 Au 293 275 7.21      
13 Au 148 139 1.59      
14 Bg 3228 3031 0.00      
15 Bg 1522 1430 0.00      
16 Bg 1060 996 0.00      
17 Bg 208 195 0.00      
18 Bu 3230 3034 24.24      
19 Bu 3114 2924 41.30      
20 Bu 1529 1436 25.81      
21 Bu 1457 1368 2.43      
22 Bu 1162 1092 2.02      
23 Bu 1064 999 5.69      
24 Bu 359 337 15.08      

Unscaled Zero Point Vibrational Energy (zpe) 18987.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 17833.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
1.37565 0.14822 0.14078

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.378 0.504 0.000
N2 -0.378 -0.504 0.000
C3 -0.378 1.758 0.000
C4 0.378 -1.758 0.000
H5 -1.453 1.580 0.000
H6 1.453 -1.580 0.000
H7 -0.078 2.323 0.880
H8 -0.078 2.323 -0.880
H9 0.078 -2.323 0.880
H10 0.078 -2.323 -0.880

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10
N11.25941.46442.26122.12362.34402.07212.07212.97582.9758
N21.25942.26121.46442.34402.12362.97582.97582.07212.0721
C31.46442.26123.59561.08923.80641.08831.08834.19954.1995
C42.26121.46443.59563.80641.08924.19954.19951.08831.0883
H52.12362.34401.08923.80644.29211.79371.79374.28374.2837
H62.34402.12363.80641.08924.29214.28374.28371.79371.7937
H72.07212.97581.08834.19951.79374.28371.75994.64924.9711
H82.07212.97581.08834.19951.79374.28371.75994.97114.6492
H92.97582.07214.19951.08834.28371.79374.64924.97111.7599
H102.97582.07214.19951.08834.28371.79374.97114.64921.7599

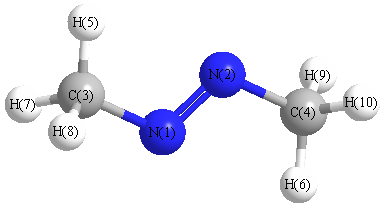
picture of (E)-1,2-Dimethyldiazene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 112.011 N1 C3 H5 111.686
N1 C3 H7 107.619 N1 C3 H8 107.619
N2 N1 C3 112.011 N2 C4 H6 111.686
N2 C4 H9 107.619 N2 C4 H10 107.619
H5 C3 H7 110.920 H5 C3 H8 110.920
H6 C4 H9 110.920 H6 C4 H10 110.920
H7 C3 H8 107.908 H9 C4 H10 107.908
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability