return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCHCHCH3 (2-Butenal)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-230.570661
Energy at 298.15K-230.576461
HF Energy-229.818515
Nuclear repulsion energy152.816339
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3262 3064 3.66      
2 A' 3232 3035 19.83      
3 A' 3212 3017 4.00      
4 A' 3116 2926 13.82      
5 A' 3005 2822 87.45      
6 A' 1750 1644 221.63      
7 A' 1713 1609 21.61      
8 A' 1534 1441 16.11      
9 A' 1458 1369 3.38      
10 A' 1444 1356 0.45      
11 A' 1352 1270 2.60      
12 A' 1309 1229 0.22      
13 A' 1203 1130 49.97      
14 A' 1130 1061 15.72      
15 A' 966 907 19.74      
16 A' 545 512 7.68      
17 A' 466 438 2.41      
18 A' 209 196 7.34      
19 A" 3198 3003 8.86      
20 A" 1520 1427 8.83      
21 A" 1086 1020 2.48      
22 A" 1024 962 18.18      
23 A" 1001 940 30.12      
24 A" 777 730 0.22      
25 A" 282 265 9.14      
26 A" 190 178 0.55      
27 A" 120 113 2.02      

Unscaled Zero Point Vibrational Energy (zpe) 20051.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 18832.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
1.09327 0.07256 0.06890

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.383 -0.485 0.000
C2 0.901 -0.664 0.000
C3 0.000 0.336 0.000
C4 -1.436 0.047 0.000
O5 -2.315 0.909 0.000
H6 -1.704 -1.025 0.000
H7 0.297 1.378 0.000
H8 0.531 -1.686 0.000
H9 2.654 0.568 0.000
H10 2.825 -0.959 0.877
H11 2.825 -0.959 -0.877

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.49222.52043.85584.90004.12262.79682.20701.08761.09021.0902
C21.49221.34612.44313.58002.63052.12921.08702.14202.13492.1349
C32.52041.34611.46482.38442.18091.08342.09022.66383.22923.2292
C43.85582.44311.46481.23071.10502.18502.62144.12284.46554.4655
O54.90003.58002.38441.23072.02792.65333.85094.97995.53865.5386
H64.12262.63052.18091.10502.02793.12692.33084.63994.61424.6142
H72.79682.12921.08342.18502.65333.12693.07252.49193.55253.5525
H82.20701.08702.09022.62143.85092.33083.07253.09612.56172.5617
H91.08762.14202.66384.12284.97994.63992.49193.09611.76881.7688
H101.09022.13493.22924.46555.53864.61423.55252.56171.76881.7534
H111.09022.13493.22924.46555.53864.61423.55252.56171.76881.7534

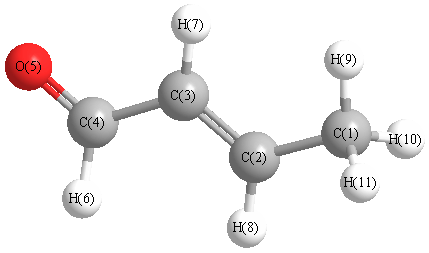
picture of 2-Butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.162 C1 C2 H8 116.802
C2 C1 H9 111.292 C2 C1 H10 110.557
C2 C1 H11 110.557 C2 C3 C4 120.662
C2 C3 H7 122.055 C3 C2 H8 118.036
C3 C4 O5 124.171 C3 C4 H6 115.420
C4 C3 H7 117.283 O5 C4 H6 120.409
H9 C1 H10 108.625 H9 C1 H11 108.625
H10 C1 H11 107.058
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability