Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -376.629603 |
| Energy at 298.15K | |
| HF Energy | -375.808003 |
| Nuclear repulsion energy | 189.849739 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3257 |
3059 |
12.54 |
33.66 |
0.71 |
0.83 |
| 2 |
A |
3220 |
3025 |
16.91 |
86.96 |
0.30 |
0.46 |
| 3 |
A |
3173 |
2980 |
13.47 |
98.80 |
0.08 |
0.15 |
| 4 |
A |
1547 |
1453 |
5.68 |
5.71 |
0.74 |
0.85 |
| 5 |
A |
1492 |
1401 |
12.96 |
1.85 |
0.67 |
0.80 |
| 6 |
A |
1425 |
1338 |
21.67 |
3.50 |
0.74 |
0.85 |
| 7 |
A |
1365 |
1282 |
18.39 |
2.38 |
0.72 |
0.83 |
| 8 |
A |
1284 |
1206 |
10.76 |
6.14 |
0.71 |
0.83 |
| 9 |
A |
1183 |
1111 |
61.08 |
4.95 |
0.72 |
0.84 |
| 10 |
A |
1158 |
1088 |
36.27 |
2.12 |
0.21 |
0.34 |
| 11 |
A |
1111 |
1044 |
251.96 |
1.27 |
0.46 |
0.63 |
| 12 |
A |
1087 |
1021 |
38.19 |
7.86 |
0.38 |
0.55 |
| 13 |
A |
928 |
872 |
43.92 |
5.00 |
0.29 |
0.44 |
| 14 |
A |
570 |
535 |
4.46 |
2.00 |
0.30 |
0.46 |
| 15 |
A |
483 |
454 |
22.11 |
0.67 |
0.71 |
0.83 |
| 16 |
A |
425 |
399 |
6.37 |
1.49 |
0.50 |
0.67 |
| 17 |
A |
246 |
231 |
10.25 |
0.06 |
0.54 |
0.70 |
| 18 |
A |
125 |
118 |
9.97 |
0.05 |
0.64 |
0.78 |
Unscaled Zero Point Vibrational Energy (zpe) 12039.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 11307.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.760 |
-0.603 |
-0.287 |
| C2 |
0.461 |
0.021 |
0.336 |
| F3 |
-1.885 |
0.100 |
0.151 |
| F4 |
1.547 |
-0.756 |
-0.008 |
| F5 |
0.663 |
1.274 |
-0.183 |
| H6 |
-0.699 |
-0.531 |
-1.370 |
| H7 |
-0.854 |
-1.639 |
0.030 |
| H8 |
0.420 |
0.104 |
1.419 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5067 | 1.3972 | 2.3297 | 2.3578 | 1.0871 | 1.0871 | 2.1920 |
C2 | 1.5067 | | 2.3555 | 1.3786 | 1.3712 | 2.1356 | 2.1395 | 1.0869 | F3 | 1.3972 | 2.3555 | | 3.5416 | 2.8256 | 2.0297 | 2.0251 | 2.6312 | F4 | 2.3297 | 1.3786 | 3.5416 | | 2.2207 | 2.6367 | 2.5591 | 2.0116 | F5 | 2.3578 | 1.3712 | 2.8256 | 2.2207 | | 2.5534 | 3.2907 | 1.9980 | H6 | 1.0871 | 2.1356 | 2.0297 | 2.6367 | 2.5534 | | 1.7921 | 3.0715 | H7 | 1.0871 | 2.1395 | 2.0251 | 2.5591 | 3.2907 | 1.7921 | | 2.5673 | H8 | 2.1920 | 1.0869 | 2.6312 | 2.0116 | 1.9980 | 3.0715 | 2.5673 | |
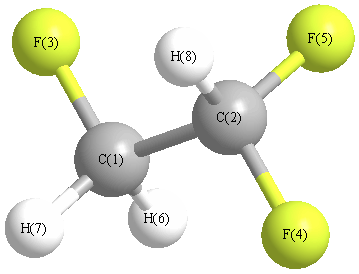 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.611 |
|
C1 |
C2 |
F5 |
109.938 |
| C1 |
C2 |
H8 |
114.407 |
|
C2 |
C1 |
F3 |
108.357 |
| C2 |
C1 |
H6 |
109.791 |
|
C2 |
C1 |
H7 |
110.099 |
| F3 |
C1 |
H6 |
108.937 |
|
F3 |
C1 |
H7 |
108.564 |
| F4 |
C2 |
F5 |
107.722 |
|
F4 |
C2 |
H8 |
108.774 |
| F5 |
C2 |
H8 |
108.190 |
|
H6 |
C1 |
H7 |
111.033 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -376.627131 |
| Energy at 298.15K | |
| HF Energy | -375.805122 |
| Nuclear repulsion energy | 192.776048 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3187 |
2994 |
38.44 |
98.67 |
0.13 |
0.24 |
| 2 |
A' |
3160 |
2968 |
10.16 |
81.39 |
0.16 |
0.27 |
| 3 |
A' |
1544 |
1450 |
6.55 |
5.36 |
0.75 |
0.86 |
| 4 |
A' |
1471 |
1382 |
21.07 |
3.04 |
0.57 |
0.72 |
| 5 |
A' |
1444 |
1356 |
24.70 |
1.95 |
0.75 |
0.86 |
| 6 |
A' |
1199 |
1126 |
104.29 |
5.98 |
0.25 |
0.40 |
| 7 |
A' |
1107 |
1039 |
22.88 |
3.86 |
0.75 |
0.86 |
| 8 |
A' |
899 |
844 |
42.04 |
7.81 |
0.10 |
0.18 |
| 9 |
A' |
759 |
713 |
55.36 |
2.50 |
0.53 |
0.69 |
| 10 |
A' |
505 |
475 |
13.09 |
1.40 |
0.74 |
0.85 |
| 11 |
A' |
238 |
223 |
1.89 |
0.17 |
0.47 |
0.64 |
| 12 |
A" |
3237 |
3040 |
10.43 |
49.46 |
0.75 |
0.86 |
| 13 |
A" |
1417 |
1331 |
30.16 |
1.52 |
0.75 |
0.86 |
| 14 |
A" |
1300 |
1221 |
15.74 |
10.62 |
0.75 |
0.86 |
| 15 |
A" |
1152 |
1082 |
113.45 |
1.31 |
0.75 |
0.86 |
| 16 |
A" |
974 |
915 |
82.53 |
3.47 |
0.75 |
0.86 |
| 17 |
A" |
378 |
355 |
0.30 |
0.22 |
0.75 |
0.86 |
| 18 |
A" |
126 |
118 |
2.39 |
0.09 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12047.4 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 11314.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.752 |
-0.825 |
0.000 |
| C2 |
0.365 |
0.630 |
0.000 |
| F3 |
-0.392 |
-1.615 |
0.000 |
| F4 |
-0.392 |
0.918 |
1.108 |
| F5 |
-0.392 |
0.918 |
-1.108 |
| H6 |
1.329 |
-1.049 |
-0.895 |
| H7 |
1.329 |
-1.049 |
0.895 |
| H8 |
1.232 |
1.290 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5055 | 1.3907 | 2.3611 | 2.3611 | 1.0883 | 1.0883 | 2.1686 |
C2 | 1.5055 | | 2.3695 | 1.3728 | 1.3728 | 2.1332 | 2.1332 | 1.0896 | F3 | 1.3907 | 2.3695 | | 2.7649 | 2.7649 | 2.0211 | 2.0211 | 3.3284 | F4 | 2.3611 | 1.3728 | 2.7649 | | 2.2166 | 3.2933 | 2.6227 | 2.0012 | F5 | 2.3611 | 1.3728 | 2.7649 | 2.2166 | | 2.6227 | 3.2933 | 2.0012 | H6 | 1.0883 | 2.1332 | 2.0211 | 3.2933 | 2.6227 | | 1.7898 | 2.5064 | H7 | 1.0883 | 2.1332 | 2.0211 | 2.6227 | 3.2933 | 1.7898 | | 2.5064 | H8 | 2.1686 | 1.0896 | 3.3284 | 2.0012 | 2.0012 | 2.5064 | 2.5064 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.148 |
|
C1 |
C2 |
F5 |
110.148 |
| C1 |
C2 |
H8 |
112.378 |
|
C2 |
C1 |
F3 |
109.733 |
| C2 |
C1 |
H6 |
109.607 |
|
C2 |
C1 |
H7 |
109.607 |
| F3 |
C1 |
H6 |
108.617 |
|
F3 |
C1 |
H7 |
108.617 |
| F4 |
C2 |
F5 |
107.674 |
|
F4 |
C2 |
H8 |
108.174 |
| F5 |
C2 |
H8 |
108.174 |
|
H6 |
C1 |
H7 |
110.636 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
