return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11 (2-Methylbut-2-yl radical)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-196.464012
Energy at 298.15K-196.475037
HF Energy-195.726504
Nuclear repulsion energy182.696211
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3223 3027 26.89      
2 A 3214 3019 19.72      
3 A 3213 3018 43.94      
4 A 3207 3012 18.23      
5 A 3163 2971 22.57      
6 A 3154 2962 26.09      
7 A 3136 2946 21.96      
8 A 3121 2931 27.54      
9 A 3047 2862 52.76      
10 A 3043 2858 33.27      
11 A 3019 2835 43.98      
12 A 1557 1463 1.69      
13 A 1548 1453 7.39      
14 A 1538 1445 6.04      
15 A 1537 1443 19.16      
16 A 1526 1433 2.35      
17 A 1522 1429 3.44      
18 A 1512 1420 0.02      
19 A 1458 1369 6.00      
20 A 1455 1367 1.65      
21 A 1444 1356 6.50      
22 A 1419 1333 0.21      
23 A 1347 1265 1.02      
24 A 1312 1232 4.33      
25 A 1284 1206 4.73      
26 A 1153 1083 2.41      
27 A 1105 1038 0.06      
28 A 1077 1011 3.60      
29 A 1019 957 0.26      
30 A 1013 951 0.92      
31 A 971 912 0.77      
32 A 960 901 0.32      
33 A 789 741 0.67      
34 A 767 721 2.07      
35 A 464 436 0.47      
36 A 387 363 0.13      
37 A 324 305 1.94      
38 A 290 272 0.01      
39 A 224 211 0.69      
40 A 159 149 0.00      
41 A 149 140 0.08      
42 A 73 69 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 32960.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 30956.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.25007 0.11547 0.08554

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.590 1.472 0.067
H2 0.266 2.005 -0.344
H3 -1.492 1.883 -0.384
H4 -0.628 1.707 1.140
C5 2.052 -0.043 -0.161
H6 2.180 0.917 0.337
H7 2.906 -0.666 0.101
H8 2.076 0.128 -1.237
C9 0.748 -0.721 0.249
H10 0.721 -1.732 -0.164
H11 0.747 -0.850 1.344
C12 -0.502 -0.003 -0.157
C13 -1.778 -0.779 -0.096
H14 -1.653 -1.784 -0.497
H15 -2.127 -0.886 0.940
H16 -2.574 -0.282 -0.650

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15 H16
C11.08861.08931.09923.05412.83774.09783.25772.57563.46962.96831.49432.55023.47162.94762.7443
H21.08861.76261.75822.72382.30513.78212.75642.83173.76913.35192.15793.46224.25013.96693.6595
H31.08931.76261.76094.04003.86505.10644.06693.49304.24433.93312.14172.69223.67223.13442.4352
H41.09921.75821.76093.45523.02544.38153.93122.92963.91762.91062.14973.00483.99013.00223.3093
C53.05412.72384.04003.45521.08821.08901.08941.52682.15112.14982.55513.90084.10814.40384.6587
H62.83772.30513.86503.02541.08821.75721.76352.17733.06522.48782.87814.32714.76254.70795.0014
H74.09783.78215.10644.38151.08901.75721.76352.16352.44542.49753.48164.68894.73185.10705.5446
H83.25772.75644.06693.93121.08941.76351.76352.16652.53883.06312.79834.11984.25524.84084.7049
C92.57562.83173.49302.92961.52682.17732.16352.16651.09251.10181.49822.54982.72992.96143.4698
H103.46963.76914.24433.91762.15113.06522.44542.53881.09251.74712.11822.67532.39783.16963.6326
H112.96833.35193.93312.91062.14982.48782.49753.06311.10181.74712.12872.90733.16582.90253.9152
C121.49432.15792.14172.14972.55512.87813.48162.79831.49822.11822.12871.49412.14782.15042.1481
C132.55023.46222.69223.00483.90084.32714.68894.11982.54982.67532.90731.49411.08981.09891.0898
H143.47164.25013.67223.99014.10814.76254.73184.25522.72992.39783.16582.14781.08981.76011.7683
H152.94763.96693.13443.00224.40384.70795.10704.84082.96143.16962.90252.15041.09891.76011.7590
H162.74433.65952.43523.30934.65875.00145.54464.70493.46983.63263.91522.14811.08981.76831.7590

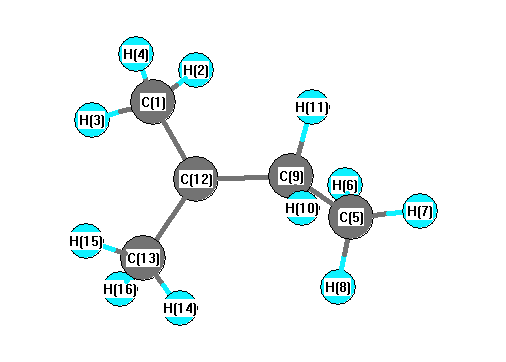
picture of 2-Methylbut-2-yl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 118.784 C1 C12 C13 117.161
H2 C1 H3 108.061 H2 C1 H4 106.951
H2 C1 C12 112.374 H3 C1 H4 107.143
H3 C1 C12 111.017 H4 C1 C12 111.052
C5 C9 H10 109.302 C5 C9 H11 108.659
C5 C9 C12 115.268 H6 C5 H7 107.620
H6 C5 H8 108.152 H6 C5 C9 111.643
H7 C5 H8 108.103 H7 C5 C9 110.490
H8 C5 C9 110.700 C9 C12 C13 116.889
H10 C9 H11 105.533 H10 C9 C12 108.682
H11 C9 C12 108.955 C12 C13 H14 111.495
C12 C13 H15 111.143 C12 C13 H16 111.516
H14 C13 H15 107.059 H14 C13 H16 108.445
H15 C13 H16 106.961
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability