return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-330.573821
Energy at 298.15K-330.579068
HF Energy-329.574052
Nuclear repulsion energy268.644951
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3291 3091 0.34      
2 A1 3280 3081 12.99      
3 A1 3259 3061 0.07      
4 A1 1667 1566 47.04      
5 A1 1548 1454 75.40      
6 A1 1256 1180 95.06      
7 A1 1197 1124 16.00      
8 A1 1053 989 6.77      
9 A1 1024 962 0.31      
10 A1 823 773 27.55      
11 A1 521 489 5.01      
12 A2 910 855 0.00      
13 A2 816 766 0.00      
14 A2 398 373 0.00      
15 B1 897 843 0.72      
16 B1 859 807 3.06      
17 B1 743 698 115.64      
18 B1 495 465 9.03      
19 B1 383 359 0.89      
20 B1 225 211 0.03      
21 B2 3289 3089 3.05      
22 B2 3268 3069 8.25      
23 B2 1676 1574 6.02      
24 B2 1507 1415 1.00      
25 B2 1471 1382 0.18      
26 B2 1348 1266 0.59      
27 B2 1202 1129 0.11      
28 B2 1107 1040 7.63      
29 B2 624 586 0.15      
30 B2 402 377 2.52      

Unscaled Zero Point Vibrational Energy (zpe) 20269.0 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 19036.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.18868 0.08511 0.05865

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.290
C2 0.000 0.000 0.923
C3 0.000 1.220 0.260
C4 0.000 -1.220 0.260
C5 0.000 1.209 -1.137
C6 0.000 -1.209 -1.137
C7 0.000 0.000 -1.836
H8 0.000 2.140 0.827
H9 0.000 -2.140 0.827
H10 0.000 2.148 -1.675
H11 0.000 -2.148 -1.675
H12 0.000 0.000 -2.918

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.36662.36792.36793.63383.63384.12622.59202.59204.50894.50895.2076
C21.36661.38831.38832.38882.38882.75972.14202.14203.37093.37093.8411
C32.36791.38832.43951.39702.80222.42561.08073.40712.14603.88403.4041
C42.36791.38832.43952.80221.39702.42563.40711.08073.88402.14603.4041
C53.63382.38881.39702.80222.41891.39722.17313.88261.08183.40032.1529
C63.63382.38882.80221.39702.41891.39723.88262.17313.40031.08182.1529
C74.12622.75972.42562.42561.39721.39723.41663.41662.15422.15421.0814
H82.59202.14201.08073.40712.17313.88263.41664.27972.50164.96434.3131
H92.59202.14203.40711.08073.88262.17313.41664.27974.96432.50164.3131
H104.50893.37092.14603.88401.08183.40032.15422.50164.96434.29612.4819
H114.50893.37093.88402.14603.40031.08182.15424.96432.50164.29612.4819
H125.20763.84113.40413.40412.15292.15291.08144.31314.31312.48192.4819

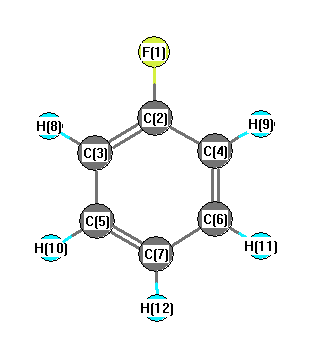
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 118.528 F1 C2 C4 118.528
C2 C3 C5 118.107 C2 C3 H8 119.837
C2 C4 C6 118.107 C2 C4 H9 119.837
C3 C2 C4 122.943 C3 C5 C7 120.469
C3 C5 H10 119.389 C4 C6 C7 120.469
C4 C6 H11 119.389 C5 C3 H8 122.056
C5 C7 C6 119.904 C5 C7 H12 120.048
C6 C4 H9 122.056 C6 C7 H12 120.048
C7 C5 H10 120.142 C7 C6 H11 120.142
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability