return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-306.812973
Energy at 298.15K-306.824340
Nuclear repulsion energy264.965387
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3249 3051 21.67      
2 A' 3213 3017 24.08      
3 A' 3198 3004 16.86      
4 A' 3128 2938 19.97      
5 A' 3070 2884 97.12      
6 A' 3029 2844 73.22      
7 A' 1559 1464 4.25      
8 A' 1545 1451 0.20      
9 A' 1513 1421 5.85      
10 A' 1440 1352 18.88      
11 A' 1341 1260 5.11      
12 A' 1229 1154 54.85      
13 A' 1205 1132 120.27      
14 A' 1130 1062 20.04      
15 A' 1019 957 36.49      
16 A' 943 886 8.41      
17 A' 861 809 16.28      
18 A' 655 615 5.37      
19 A' 491 461 0.98      
20 A' 449 421 10.46      
21 A' 275 258 2.58      
22 A" 3210 3015 30.46      
23 A" 3065 2878 17.43      
24 A" 1542 1449 3.14      
25 A" 1463 1374 17.40      
26 A" 1420 1334 1.05      
27 A" 1398 1313 0.08      
28 A" 1354 1272 1.98      
29 A" 1281 1203 29.51      
30 A" 1250 1174 0.76      
31 A" 1111 1044 47.48      
32 A" 1053 989 90.76      
33 A" 932 875 21.28      
34 A" 914 859 12.06      
35 A" 459 432 7.86      
36 A" 273 257 2.75      

Unscaled Zero Point Vibrational Energy (zpe) 27633.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 25953.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.16700 0.16038 0.09207

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.630 -1.202 0.000
O2 0.017 -0.766 1.175
O3 0.017 -0.766 -1.175
C4 0.017 0.665 1.236
C5 0.017 0.665 -1.236
C6 0.691 1.238 0.000
H7 -0.592 -2.286 0.000
H8 -1.669 -0.838 0.000
H9 0.540 0.927 2.153
H10 -1.018 1.024 1.307
H11 0.540 0.927 -2.153
H12 -1.018 1.024 -1.307
H13 1.743 0.954 0.000
H14 0.625 2.328 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.41061.41062.33112.33112.77541.08411.10103.24612.61073.24612.61073.20643.7474
O21.41062.34991.43222.80392.41902.01532.05642.02372.07193.77003.23042.70503.3653
O31.41062.34992.80391.43222.41902.01532.05643.77003.23042.02372.07192.70503.3653
C42.33111.43222.80392.47271.52053.25672.57501.08701.09793.43922.76932.14272.1600
C52.33112.80391.43222.47271.52053.25672.57503.43922.76931.08701.09792.14272.1600
C62.77542.41902.41901.52051.52053.75043.14362.18052.16242.18052.16241.08961.0922
H71.08412.01532.01533.25673.25673.75041.80454.02953.58404.02953.58403.99344.7721
H81.10102.05642.05642.57502.57503.14361.80453.55352.36633.55352.36633.85403.9103
H93.24612.02373.77001.08703.43922.18054.02953.55351.77494.30573.79542.46662.5702
H102.61072.07193.23041.09792.76932.16243.58402.36631.77493.79542.61403.05562.4718
H113.24613.77002.02373.43921.08702.18054.02953.55354.30573.79541.77492.46662.5702
H122.61073.23042.07192.76931.09792.16243.58402.36633.79542.61401.77493.05562.4718
H133.20642.70502.70502.14272.14271.08963.99343.85402.46663.05562.46663.05561.7718
H143.74743.36533.36532.16002.16001.09224.77213.91032.57022.47182.57022.47181.7718

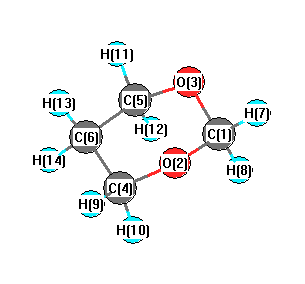
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.170 C1 O3 C5 110.170
O2 C1 O3 112.799 O2 C1 H7 107.040
O2 C1 H8 109.304 O2 C4 C6 109.991
O2 C4 H9 106.086 O2 C4 H10 109.243
O3 C1 H7 107.040 O3 C1 H8 109.304
O3 C5 C6 109.991 O3 C5 H11 106.086
O3 C5 H12 109.243 C4 C6 C5 108.807
C4 C6 H13 109.243 C4 C6 H14 110.463
C5 C6 H13 109.243 C5 C6 H14 110.463
C6 C4 H9 112.432 C6 C4 H10 110.306
C6 C5 H11 112.432 C6 C5 H12 110.306
H7 C1 H8 111.345 H9 C4 H10 108.657
H11 C5 H12 108.657 H13 C6 H14 108.600
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability