Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -132.893278 |
| Energy at 298.15K | -132.897496 |
| HF Energy | -132.479168 |
| Nuclear repulsion energy | 65.250750 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3571 |
3354 |
0.67 |
|
|
|
| 2 |
A' |
3393 |
3187 |
1.38 |
|
|
|
| 3 |
A' |
3268 |
3069 |
4.61 |
|
|
|
| 4 |
A' |
3163 |
2971 |
38.40 |
|
|
|
| 5 |
A' |
1556 |
1461 |
10.94 |
|
|
|
| 6 |
A' |
1502 |
1411 |
9.16 |
|
|
|
| 7 |
A' |
1411 |
1325 |
13.07 |
|
|
|
| 8 |
A' |
1257 |
1180 |
26.84 |
|
|
|
| 9 |
A' |
1086 |
1020 |
10.35 |
|
|
|
| 10 |
A' |
1033 |
970 |
10.44 |
|
|
|
| 11 |
A' |
504 |
474 |
16.47 |
|
|
|
| 12 |
A" |
1136 |
1067 |
0.01 |
|
|
|
| 13 |
A" |
805 |
756 |
1.54 |
|
|
|
| 14 |
A" |
725 |
681 |
146.02 |
|
|
|
| 15 |
A" |
524 |
492 |
2.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12466.2 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 11708.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.417 |
0.000 |
| C2 |
1.143 |
-0.396 |
0.000 |
| N3 |
-1.182 |
-0.115 |
0.000 |
| H4 |
0.153 |
1.497 |
0.000 |
| H5 |
2.133 |
0.029 |
0.000 |
| H6 |
1.025 |
-1.467 |
0.000 |
| H7 |
-1.895 |
0.612 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4026 | 1.2961 | 1.0907 | 2.1682 | 2.1453 | 1.9054 |
C2 | 1.4026 | | 2.3418 | 2.1363 | 1.0774 | 1.0780 | 3.2012 | N3 | 1.2961 | 2.3418 | | 2.0928 | 3.3182 | 2.5884 | 1.0186 | H4 | 1.0907 | 2.1363 | 2.0928 | | 2.4653 | 3.0902 | 2.2312 | H5 | 2.1682 | 1.0774 | 3.3182 | 2.4653 | | 1.8620 | 4.0705 | H6 | 2.1453 | 1.0780 | 2.5884 | 3.0902 | 1.8620 | | 3.5852 | H7 | 1.9054 | 3.2012 | 1.0186 | 2.2312 | 4.0705 | 3.5852 | |
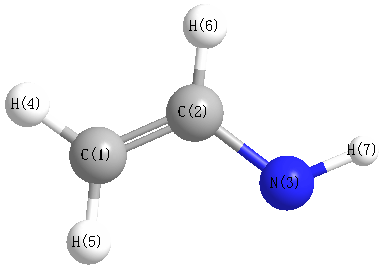 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.356 |
|
C1 |
C2 |
H6 |
119.142 |
| C1 |
N3 |
H7 |
110.228 |
|
C2 |
C1 |
N3 |
120.342 |
| C2 |
C1 |
H4 |
117.374 |
|
N3 |
C1 |
H4 |
122.284 |
| H5 |
C2 |
H6 |
119.502 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -132.891620 |
| Energy at 298.15K | -132.895822 |
| HF Energy | -132.478019 |
| Nuclear repulsion energy | 65.231632 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3519 |
3305 |
3.29 |
|
|
|
| 2 |
A' |
3375 |
3170 |
3.69 |
|
|
|
| 3 |
A' |
3254 |
3056 |
3.79 |
|
|
|
| 4 |
A' |
3232 |
3036 |
17.03 |
|
|
|
| 5 |
A' |
1555 |
1460 |
6.30 |
|
|
|
| 6 |
A' |
1484 |
1394 |
1.16 |
|
|
|
| 7 |
A' |
1409 |
1324 |
17.43 |
|
|
|
| 8 |
A' |
1246 |
1170 |
33.41 |
|
|
|
| 9 |
A' |
1107 |
1040 |
36.45 |
|
|
|
| 10 |
A' |
1017 |
956 |
0.02 |
|
|
|
| 11 |
A' |
506 |
475 |
10.40 |
|
|
|
| 12 |
A" |
1162 |
1092 |
61.39 |
|
|
|
| 13 |
A" |
817 |
767 |
84.46 |
|
|
|
| 14 |
A" |
718 |
675 |
0.06 |
|
|
|
| 15 |
A" |
499 |
469 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12450.1 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 11693.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.119 |
-0.398 |
0.000 |
| N3 |
-1.245 |
0.083 |
0.000 |
| H4 |
0.181 |
1.520 |
0.000 |
| H5 |
2.121 |
0.000 |
0.000 |
| H6 |
0.996 |
-1.471 |
0.000 |
| H7 |
-1.299 |
-0.938 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4038 | 1.2978 | 1.0857 | 2.1677 | 2.1633 | 1.9011 |
C2 | 1.4038 | | 2.4124 | 2.1350 | 1.0778 | 1.0799 | 2.4782 | N3 | 1.2978 | 2.4124 | | 2.0248 | 3.3665 | 2.7267 | 1.0223 | H4 | 1.0857 | 2.1350 | 2.0248 | | 2.4638 | 3.0998 | 2.8697 | H5 | 2.1677 | 1.0778 | 3.3665 | 2.4638 | | 1.8519 | 3.5466 | H6 | 2.1633 | 1.0799 | 2.7267 | 3.0998 | 1.8519 | | 2.3565 | H7 | 1.9011 | 2.4782 | 1.0223 | 2.8697 | 3.5466 | 2.3565 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.174 |
|
C1 |
C2 |
H6 |
120.589 |
| C1 |
N3 |
H7 |
109.479 |
|
C2 |
C1 |
N3 |
126.447 |
| C2 |
C1 |
H4 |
117.523 |
|
N3 |
C1 |
H4 |
116.030 |
| H5 |
C2 |
H6 |
118.237 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
