return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Butene, 2-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-195.910359
Energy at 298.15K-195.920031
HF Energy-195.164412
Nuclear repulsion energy174.566519
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3244 3047 35.79      
2 A 3230 3034 21.45      
3 A 3214 3019 28.97      
4 A 3208 3013 5.19      
5 A 3113 2924 24.93      
6 A 3107 2918 44.38      
7 A 3098 2910 31.72      
8 A 1758 1651 0.96      
9 A 1547 1453 10.92      
10 A 1545 1451 5.22      
11 A 1531 1438 2.22      
12 A 1466 1376 1.33      
13 A 1456 1367 1.64      
14 A 1453 1365 7.89      
15 A 1401 1316 2.08      
16 A 1271 1194 3.43      
17 A 1164 1094 6.43      
18 A 1094 1028 4.16      
19 A 996 936 1.96      
20 A 988 928 1.12      
21 A 788 740 1.31      
22 A 526 494 1.62      
23 A 388 364 0.16      
24 A 302 284 0.18      
25 A 3185 2991 16.50      
26 A 3178 2985 23.65      
27 A 3175 2982 17.39      
28 A 1531 1438 21.14      
29 A 1523 1431 0.20      
30 A 1512 1420 0.99      
31 A 1122 1054 1.56      
32 A 1072 1007 1.19      
33 A 1031 969 3.09      
34 A 820 770 16.67      
35 A 415 390 3.31      
36 A 262 246 1.92      
37 A 175 164 1.60      
38 A 107 100 0.65      
39 A 81 76 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 30538.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 28682.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.26945 0.11728 0.08557

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.502 0.498 0.000
H2 -1.961 -0.486 0.000
H3 -1.862 1.040 0.877
H4 -1.862 1.040 -0.877
C5 0.261 -2.094 0.000
H6 0.635 -2.625 -0.877
H7 0.635 -2.625 0.877
H8 -0.822 -2.173 0.000
C9 0.739 -0.674 0.000
H10 1.819 -0.553 0.000
C11 0.000 0.452 0.000
C12 0.660 1.803 0.000
H13 1.746 1.719 0.000
H14 0.360 2.380 0.877
H15 0.360 2.380 -0.877

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 C12 H13 H14 H15
C11.08591.09161.09163.13503.88403.88402.75592.52953.48351.50302.52623.46972.78962.7896
H21.08591.76281.76282.74323.47593.47592.03532.70743.78092.17423.48084.31323.79163.7916
H31.09161.76281.75343.88554.76834.43413.48873.23604.10522.14022.77713.77392.59503.1322
H41.09161.76281.75343.88554.43414.76833.48873.23604.10522.14022.77713.77393.13222.5950
C53.13502.74323.88553.88551.09081.09081.08631.49862.19142.55943.91804.09184.56054.5605
H63.88403.47594.76834.43411.09081.75361.75962.14122.54223.26144.51414.56825.31035.0122
H73.88403.47594.43414.76831.09081.75361.75962.14122.54223.26144.51414.56825.01225.3103
H82.75592.03533.48873.48871.08631.75961.75962.16473.09852.75064.24384.66264.78524.7852
C92.52952.70743.23603.23601.49862.14122.14122.16471.08621.34702.47872.59573.20013.2001
H103.48353.78094.10524.10522.19142.54222.54223.09851.08622.07802.62582.27303.39113.3911
C111.50302.17422.14022.14022.55943.26143.26142.75061.34702.07801.50422.15692.14882.1488
C122.52623.48082.77712.77713.91804.51414.51414.24382.47872.62581.50421.08851.09181.0918
H133.46974.31323.77393.77394.09184.56824.56824.66262.59572.27302.15691.08851.76781.7678
H142.78963.79162.59503.13224.56055.31035.01224.78523.20013.39112.14881.09181.76781.7545
H152.78963.79163.13222.59504.56055.01225.31034.78523.20013.39112.14881.09181.76781.7545

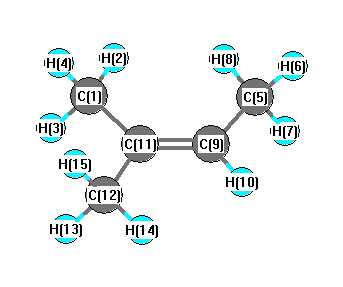
picture of 2-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C11 C9 125.047 C1 C11 C12 114.288
H2 C1 H3 108.105 H2 C1 H4 108.105
H2 C1 C11 113.262 H3 C1 H4 106.856
H3 C1 C11 110.140 H4 C1 C11 110.140
C5 C9 H10 115.010 C5 C9 C11 128.090
H6 C5 H7 106.990 H6 C5 H8 107.846
H6 C5 C9 110.581 H7 C5 H8 107.846
H7 C5 C9 110.581 H8 C5 C9 112.772
C9 C11 C12 120.665 H10 C9 C11 116.899
C11 C12 H13 111.581 C11 C12 H14 110.730
C11 C12 H15 110.730 H13 C12 H14 108.355
H13 C12 H15 108.355 H14 C12 H15 106.931
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability