return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(SH)CH2CH3 (2-Butanethiol)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-555.580917
Energy at 298.15K-555.591836
HF Energy-554.824304
Nuclear repulsion energy238.031249
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3231 3035 20.44      
2 A 3226 3030 13.89      
3 A 3209 3014 32.43      
4 A 3200 3005 28.17      
5 A 3154 2962 23.28      
6 A 3126 2936 8.69      
7 A 3119 2930 24.35      
8 A 3109 2920 29.86      
9 A 3098 2909 21.79      
10 A 2809 2638 7.71      
11 A 1554 1460 5.80      
12 A 1541 1448 10.82      
13 A 1538 1445 3.66      
14 A 1537 1443 8.06      
15 A 1525 1433 1.25      
16 A 1457 1368 10.91      
17 A 1454 1365 1.92      
18 A 1414 1328 1.29      
19 A 1365 1282 10.79      
20 A 1334 1253 0.35      
21 A 1309 1229 3.65      
22 A 1219 1145 1.13      
23 A 1162 1091 5.42      
24 A 1156 1086 6.81      
25 A 1075 1009 0.19      
26 A 1040 976 4.73      
27 A 996 936 2.26      
28 A 913 858 8.57      
29 A 888 834 2.60      
30 A 813 764 5.42      
31 A 648 609 2.83      
32 A 469 441 0.46      
33 A 392 369 1.36      
34 A 344 324 1.31      
35 A 266 250 0.59      
36 A 239 225 1.26      
37 A 225 211 0.87      
38 A 211 198 17.55      
39 A 112 105 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 29738.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 27930.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.15109 0.10208 0.06663

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.971 -1.203 -0.004
H2 1.215 -0.957 1.284
C3 1.205 1.557 -0.017
H4 1.391 1.596 1.057
H5 0.785 2.517 -0.320
C6 0.233 0.432 -0.348
H7 0.070 0.411 -1.429
C8 -1.116 0.632 0.340
H9 -1.434 1.660 0.140
H10 -0.976 0.556 1.422
C11 -2.205 -0.328 -0.118
H12 -2.371 -0.236 -1.191
H13 -1.930 -1.359 0.094
H14 -3.146 -0.113 0.386
H15 2.158 1.417 -0.522

Atom - Atom Distances (Å)
  S1 H2 C3 H4 H5 C6 H7 C8 H9 H10 C11 H12 H13 H14 H15
S11.33372.77043.02303.73861.82752.33442.80123.74212.98683.29633.67662.90744.27692.9224
H21.33372.83032.56853.85012.35693.24642.97473.89452.66573.74824.41603.38634.53113.1277
C32.77042.83031.09081.09061.52282.14372.52372.64532.79803.89724.16884.28294.67771.0876
H43.02302.56851.09081.76442.16093.05462.78052.97112.61164.24424.75034.54874.89481.7642
H53.73863.85011.09061.76442.15692.48542.75762.42343.15994.13254.27794.75094.78271.7705
C61.82752.35691.52282.16092.15691.09341.52742.12652.14672.56382.81752.84313.50042.1689
H72.33443.24642.14373.05462.48541.09342.14082.50613.03982.72722.53603.07433.72932.4888
C82.80122.97472.52372.78052.75761.52742.14081.09371.09361.52202.16152.16522.16283.4751
H93.74213.89452.64532.97112.42342.12652.50611.09371.75242.14762.49853.05942.47693.6601
H102.98682.66572.79802.61163.15992.14673.03981.09361.75242.15873.06582.51762.49563.7869
C113.29633.74823.89724.24424.13252.56382.72721.52202.14762.15871.09031.08741.08904.7157
H123.67664.41604.16884.75034.27792.81752.53602.16152.49853.06581.09031.76291.76144.8670
H132.90743.38634.28294.54874.75092.84313.07432.16523.05942.51761.08741.76291.76474.9794
H144.27694.53114.67774.89484.78273.50043.72932.16282.47692.49561.08901.76141.76475.5941
H152.92243.12771.08761.76421.77052.16892.48883.47513.66013.78694.71574.86704.97945.5941

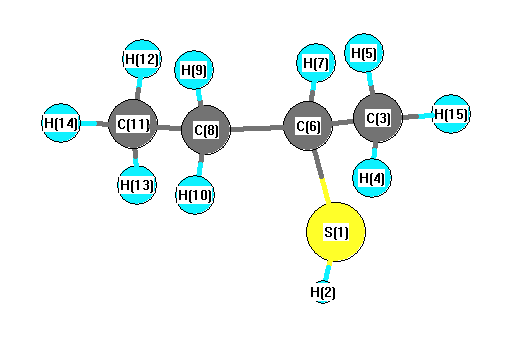
picture of 2-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C6 C3 111.246 S1 C6 H7 103.221
S1 C6 C8 112.922 H2 S1 C6 95.142
C3 C6 H7 108.942 C3 C6 C8 111.666
H4 C3 H5 107.963 H4 C3 C6 110.450
H4 C3 H15 108.164 H5 C3 C6 110.145
H5 C3 H15 108.743 C6 C3 H15 111.283
C6 C8 H9 107.293 C6 C8 H10 108.850
C6 C8 C11 114.436 H7 C6 C8 108.404
C8 C11 H12 110.580 C8 C11 H13 111.057
C8 C11 H14 110.767 H9 C8 H10 106.484
H9 C8 C11 109.285 H10 C8 C11 110.159
H12 C11 H13 108.105 H12 C11 H14 107.855
H13 C11 H14 108.364
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability