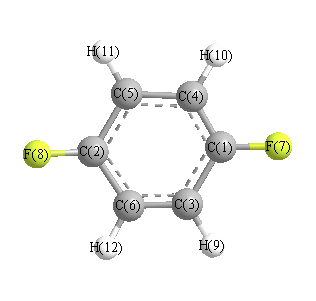
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3298 |
3097 |
0.00 |
283.33 |
0.14 |
0.25 |
| 2 |
Ag |
1677 |
1575 |
0.00 |
5.62 |
0.74 |
0.85 |
| 3 |
Ag |
1281 |
1203 |
0.00 |
29.80 |
0.07 |
0.12 |
| 4 |
Ag |
1182 |
1110 |
0.00 |
7.85 |
0.53 |
0.70 |
| 5 |
Ag |
868 |
815 |
0.00 |
37.83 |
0.05 |
0.10 |
| 6 |
Ag |
453 |
426 |
0.00 |
5.81 |
0.38 |
0.55 |
| 7 |
Au |
897 |
843 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
408 |
383 |
0.00 |
0.00 |
0.00 |
0.00 |
| 9 |
B1g |
796 |
747 |
0.00 |
0.33 |
0.75 |
0.86 |
| 10 |
B1u |
3284 |
3084 |
2.38 |
0.00 |
0.00 |
0.00 |
| 11 |
B1u |
1562 |
1467 |
239.85 |
0.00 |
0.00 |
0.00 |
| 12 |
B1u |
1232 |
1157 |
197.47 |
0.00 |
0.00 |
0.00 |
| 13 |
B1u |
1037 |
974 |
7.25 |
0.00 |
0.00 |
0.00 |
| 14 |
B1u |
745 |
700 |
62.07 |
0.00 |
0.00 |
0.00 |
| 15 |
B2g |
854 |
802 |
0.00 |
0.11 |
0.75 |
0.86 |
| 16 |
B2g |
445 |
418 |
0.00 |
3.72 |
0.75 |
0.86 |
| 17 |
B2g |
329 |
309 |
0.00 |
0.98 |
0.75 |
0.86 |
| 18 |
B2u |
3297 |
3097 |
0.00 |
0.00 |
0.75 |
0.86 |
| 19 |
B2u |
1478 |
1388 |
0.60 |
0.00 |
0.00 |
0.00 |
| 20 |
B2u |
1448 |
1360 |
0.11 |
0.00 |
0.00 |
0.00 |
| 21 |
B2u |
1124 |
1056 |
11.60 |
0.00 |
0.00 |
0.00 |
| 22 |
B2u |
345 |
324 |
5.51 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3286 |
3086 |
0.00 |
113.35 |
0.75 |
0.86 |
| 24 |
B3g |
1683 |
1580 |
0.00 |
7.48 |
0.75 |
0.86 |
| 25 |
B3g |
1321 |
1241 |
0.00 |
1.16 |
0.75 |
0.86 |
| 26 |
B3g |
644 |
605 |
0.00 |
6.67 |
0.75 |
0.86 |
| 27 |
B3g |
441 |
414 |
0.00 |
0.07 |
0.75 |
0.86 |
| 28 |
B3u |
815 |
766 |
99.57 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
494 |
464 |
15.32 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
155 |
145 |
1.95 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 18439.5 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 17318.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.