Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -577.778728 |
| Energy at 298.15K | -577.786482 |
| Nuclear repulsion energy | 159.108624 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3221 |
3025 |
23.90 |
|
|
|
| 2 |
A' |
3162 |
2970 |
27.02 |
|
|
|
| 3 |
A' |
3138 |
2947 |
9.87 |
|
|
|
| 4 |
A' |
3119 |
2930 |
23.72 |
|
|
|
| 5 |
A' |
1554 |
1459 |
7.21 |
|
|
|
| 6 |
A' |
1540 |
1446 |
0.78 |
|
|
|
| 7 |
A' |
1534 |
1441 |
2.24 |
|
|
|
| 8 |
A' |
1461 |
1372 |
2.63 |
|
|
|
| 9 |
A' |
1428 |
1341 |
15.66 |
|
|
|
| 10 |
A' |
1339 |
1258 |
22.51 |
|
|
|
| 11 |
A' |
1157 |
1087 |
0.37 |
|
|
|
| 12 |
A' |
1088 |
1022 |
1.70 |
|
|
|
| 13 |
A' |
941 |
884 |
12.36 |
|
|
|
| 14 |
A' |
792 |
743 |
28.94 |
|
|
|
| 15 |
A' |
377 |
354 |
2.11 |
|
|
|
| 16 |
A' |
242 |
228 |
2.01 |
|
|
|
| 17 |
A" |
3236 |
3039 |
21.21 |
|
|
|
| 18 |
A" |
3214 |
3019 |
25.89 |
|
|
|
| 19 |
A" |
3191 |
2997 |
0.54 |
|
|
|
| 20 |
A" |
1547 |
1453 |
9.05 |
|
|
|
| 21 |
A" |
1352 |
1270 |
0.04 |
|
|
|
| 22 |
A" |
1287 |
1209 |
0.89 |
|
|
|
| 23 |
A" |
1142 |
1072 |
1.26 |
|
|
|
| 24 |
A" |
899 |
844 |
0.04 |
|
|
|
| 25 |
A" |
774 |
727 |
2.00 |
|
|
|
| 26 |
A" |
245 |
230 |
0.01 |
|
|
|
| 27 |
A" |
124 |
117 |
1.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21551.4 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 20241.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.581 |
0.000 |
| C2 |
0.893 |
-0.643 |
0.000 |
| C3 |
2.364 |
-0.245 |
0.000 |
| Cl4 |
-1.732 |
0.141 |
0.000 |
| H5 |
0.174 |
1.190 |
0.884 |
| H6 |
0.174 |
1.190 |
-0.884 |
| H7 |
0.667 |
-1.250 |
-0.877 |
| H8 |
0.667 |
-1.250 |
0.877 |
| H9 |
3.003 |
-1.126 |
0.000 |
| H10 |
2.611 |
0.346 |
-0.882 |
| H11 |
2.611 |
0.346 |
0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5146 | 2.5044 | 1.7874 | 1.0874 | 1.0874 | 2.1363 | 2.1363 | 3.4539 | 2.7657 | 2.7657 |
C2 | 1.5146 | | 1.5246 | 2.7393 | 2.1576 | 2.1576 | 1.0900 | 1.0900 | 2.1645 | 2.1697 | 2.1697 | C3 | 2.5044 | 1.5246 | | 4.1148 | 2.7633 | 2.7633 | 2.1590 | 2.1590 | 1.0882 | 1.0895 | 1.0895 | Cl4 | 1.7874 | 2.7393 | 4.1148 | | 2.3489 | 2.3489 | 2.9079 | 2.9079 | 4.9013 | 4.4366 | 4.4366 | H5 | 1.0874 | 2.1576 | 2.7633 | 2.3489 | | 1.7679 | 3.0484 | 2.4886 | 3.7608 | 3.1251 | 2.5786 | H6 | 1.0874 | 2.1576 | 2.7633 | 2.3489 | 1.7679 | | 2.4886 | 3.0484 | 3.7608 | 2.5786 | 3.1251 | H7 | 2.1363 | 1.0900 | 2.1590 | 2.9079 | 3.0484 | 2.4886 | | 1.7534 | 2.4981 | 2.5154 | 3.0690 | H8 | 2.1363 | 1.0900 | 2.1590 | 2.9079 | 2.4886 | 3.0484 | 1.7534 | | 2.4981 | 3.0690 | 2.5154 | H9 | 3.4539 | 2.1645 | 1.0882 | 4.9013 | 3.7608 | 3.7608 | 2.4981 | 2.4981 | | 1.7602 | 1.7602 | H10 | 2.7657 | 2.1697 | 1.0895 | 4.4366 | 3.1251 | 2.5786 | 2.5154 | 3.0690 | 1.7602 | | 1.7632 | H11 | 2.7657 | 2.1697 | 1.0895 | 4.4366 | 2.5786 | 3.1251 | 3.0690 | 2.5154 | 1.7602 | 1.7632 | |
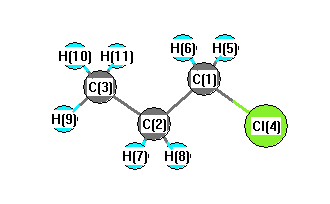 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
110.981 |
|
C1 |
C2 |
H7 |
109.121 |
| C1 |
C2 |
H8 |
109.121 |
|
C2 |
C1 |
Cl4 |
111.852 |
| C2 |
C1 |
H5 |
110.970 |
|
C2 |
C1 |
H6 |
110.970 |
| C2 |
C3 |
H9 |
110.768 |
|
C2 |
C3 |
H10 |
111.102 |
| C2 |
C3 |
H11 |
111.102 |
|
C3 |
C2 |
H7 |
110.219 |
| C3 |
C2 |
H8 |
110.219 |
|
Cl4 |
C1 |
H5 |
107.052 |
| Cl4 |
C1 |
H6 |
107.052 |
|
H5 |
C1 |
H6 |
108.761 |
| H7 |
C2 |
H8 |
107.086 |
|
H9 |
C3 |
H10 |
107.854 |
| H9 |
C3 |
H11 |
107.854 |
|
H10 |
C3 |
H11 |
108.021 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -577.778587 |
| Energy at 298.15K | -577.786453 |
| Nuclear repulsion energy | 163.242005 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3235 |
3039 |
13.95 |
|
|
|
| 2 |
A |
3234 |
3037 |
15.62 |
|
|
|
| 3 |
A |
3216 |
3020 |
30.30 |
|
|
|
| 4 |
A |
3180 |
2987 |
16.68 |
|
|
|
| 5 |
A |
3161 |
2968 |
15.10 |
|
|
|
| 6 |
A |
3124 |
2934 |
20.25 |
|
|
|
| 7 |
A |
3117 |
2928 |
23.79 |
|
|
|
| 8 |
A |
1555 |
1460 |
6.93 |
|
|
|
| 9 |
A |
1546 |
1452 |
6.62 |
|
|
|
| 10 |
A |
1527 |
1434 |
3.92 |
|
|
|
| 11 |
A |
1522 |
1430 |
5.39 |
|
|
|
| 12 |
A |
1464 |
1375 |
7.10 |
|
|
|
| 13 |
A |
1422 |
1335 |
0.42 |
|
|
|
| 14 |
A |
1390 |
1305 |
35.98 |
|
|
|
| 15 |
A |
1323 |
1243 |
1.47 |
|
|
|
| 16 |
A |
1277 |
1199 |
0.67 |
|
|
|
| 17 |
A |
1150 |
1080 |
0.48 |
|
|
|
| 18 |
A |
1125 |
1057 |
1.20 |
|
|
|
| 19 |
A |
1090 |
1023 |
2.74 |
|
|
|
| 20 |
A |
933 |
876 |
7.65 |
|
|
|
| 21 |
A |
899 |
844 |
3.79 |
|
|
|
| 22 |
A |
821 |
771 |
14.88 |
|
|
|
| 23 |
A |
707 |
664 |
13.93 |
|
|
|
| 24 |
A |
435 |
409 |
1.11 |
|
|
|
| 25 |
A |
306 |
288 |
0.35 |
|
|
|
| 26 |
A |
226 |
213 |
0.85 |
|
|
|
| 27 |
A |
138 |
130 |
0.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21560.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 20249.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.187 |
0.879 |
0.307 |
| C2 |
-1.140 |
0.565 |
-0.356 |
| C3 |
-1.785 |
-0.719 |
0.144 |
| Cl4 |
1.436 |
-0.346 |
-0.068 |
| H5 |
0.578 |
1.835 |
-0.030 |
| H6 |
0.083 |
0.903 |
1.390 |
| H7 |
-1.800 |
1.412 |
-0.153 |
| H8 |
-1.000 |
0.523 |
-1.437 |
| H9 |
-2.754 |
-0.874 |
-0.328 |
| H10 |
-1.160 |
-1.582 |
-0.075 |
| H11 |
-1.940 |
-0.678 |
1.222 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5163 | 2.5440 | 1.7896 | 1.0865 | 1.0879 | 2.1083 | 2.1392 | 3.4818 | 2.8315 | 2.7905 |
C2 | 1.5163 | | 1.5219 | 2.7477 | 2.1610 | 2.1582 | 1.0929 | 1.0906 | 2.1621 | 2.1652 | 2.1626 | C3 | 2.5440 | 1.5219 | | 3.2501 | 3.4842 | 2.7703 | 2.1521 | 2.1589 | 1.0884 | 1.0873 | 1.0900 | Cl4 | 1.7896 | 2.7477 | 3.2501 | | 2.3442 | 2.3489 | 3.6841 | 2.9263 | 4.2310 | 2.8757 | 3.6298 | H5 | 1.0865 | 2.1610 | 3.4842 | 2.3442 | | 1.7693 | 2.4184 | 2.4873 | 4.3040 | 3.8338 | 3.7717 | H6 | 1.0879 | 2.1582 | 2.7703 | 2.3489 | 1.7693 | | 2.4872 | 3.0506 | 3.7623 | 3.1409 | 2.5731 | H7 | 2.1083 | 1.0929 | 2.1521 | 3.6841 | 2.4184 | 2.4872 | | 1.7544 | 2.4828 | 3.0625 | 2.5061 | H8 | 2.1392 | 1.0906 | 2.1589 | 2.9263 | 2.4873 | 3.0506 | 1.7544 | | 2.5013 | 2.5125 | 3.0657 | H9 | 3.4818 | 2.1621 | 1.0884 | 4.2310 | 4.3040 | 3.7623 | 2.4828 | 2.5013 | | 1.7619 | 1.7619 | H10 | 2.8315 | 2.1652 | 1.0873 | 2.8757 | 3.8338 | 3.1409 | 3.0625 | 2.5125 | 1.7619 | | 1.7626 | H11 | 2.7905 | 2.1626 | 1.0900 | 3.6298 | 3.7717 | 2.5731 | 2.5061 | 3.0657 | 1.7619 | 1.7626 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.715 |
|
C1 |
C2 |
H7 |
106.673 |
| C1 |
C2 |
H8 |
109.198 |
|
C2 |
C1 |
Cl4 |
112.170 |
| C2 |
C1 |
H5 |
111.169 |
|
C2 |
C1 |
H6 |
110.861 |
| C2 |
C3 |
H9 |
110.751 |
|
C2 |
C3 |
H10 |
111.069 |
| C2 |
C3 |
H11 |
110.693 |
|
C3 |
C2 |
H7 |
109.692 |
| C3 |
C2 |
H8 |
110.363 |
|
Cl4 |
C1 |
H5 |
106.621 |
| Cl4 |
C1 |
H6 |
106.894 |
|
H5 |
C1 |
H6 |
108.920 |
| H7 |
C2 |
H8 |
106.921 |
|
H9 |
C3 |
H10 |
108.153 |
| H9 |
C3 |
H11 |
107.960 |
|
H10 |
C3 |
H11 |
108.099 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
