return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,3-dichlorobenzene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-1149.623314
Energy at 298.15K 
HF Energy-1148.513996
Nuclear repulsion energy452.647140
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3290 3090 2.92      
2 A1 3288 3089 0.10      
3 A1 3266 3067 3.70      
4 A1 1643 1544 24.02      
5 A1 1463 1374 11.76      
6 A1 1171 1100 19.92      
7 A1 1116 1048 15.27      
8 A1 1017 955 4.21      
9 A1 681 640 1.45      
10 A1 410 385 2.15      
11 A1 200 188 0.29      
12 A2 854 802 0.00      
13 A2 403 378 0.00      
14 A2 190 179 0.00      
15 B1 903 848 0.36      
16 B1 822 772 2.20      
17 B1 750 705 81.14      
18 B1 405 380 4.32      
19 B1 159 150 0.33      
20 B1 353i 331i 0.06      
21 B2 3285 3085 0.65      
22 B2 1648 1548 80.53      
23 B2 1519 1427 57.59      
24 B2 1468 1378 1.37      
25 B2 1322 1241 2.55      
26 B2 1213 1139 0.73      
27 B2 1133 1064 22.97      
28 B2 815 766 70.36      
29 B2 437 411 2.55      
30 B2 373 350 0.33      

Unscaled Zero Point Vibrational Energy (zpe) 17444.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16383.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.09419 0.02867 0.02198

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.076
C2 0.000 1.214 1.389
C3 0.000 -1.214 1.389
C4 0.000 1.199 -0.005
C5 0.000 -1.199 -0.005
C6 0.000 0.000 -0.718
Cl7 0.000 2.697 -0.881
Cl8 0.000 -2.697 -0.881
H9 0.000 0.000 3.158
H10 0.000 2.156 1.920
H11 0.000 -2.156 1.920
H12 0.000 0.000 -1.799

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.39541.39542.40222.40222.79454.00284.00281.08182.16122.16123.8750
C21.39542.42891.39482.78712.43222.71174.52282.14581.08073.41163.4113
C31.39542.42892.78711.39482.43224.52282.71172.14583.41161.08073.4113
C42.40221.39482.78712.39711.39441.73583.99303.38312.15063.86772.1568
C52.40222.78711.39482.39711.39443.99301.73583.38313.86772.15062.1568
C62.79452.43222.43221.39441.39442.70222.70223.87643.40713.40711.0805
Cl74.00282.71174.52281.73583.99302.70225.39454.85712.85355.60352.8490
Cl84.00284.52282.71173.99301.73582.70225.39454.85715.60352.85352.8490
H91.08182.14582.14583.38313.38313.87644.85714.85712.48572.48574.9569
H102.16121.08073.41162.15063.86773.40712.85355.60352.48574.31124.2986
H112.16123.41161.08073.86772.15063.40715.60352.85352.48574.31124.2986
H123.87503.41133.41132.15682.15681.08052.84902.84904.95694.29864.2986

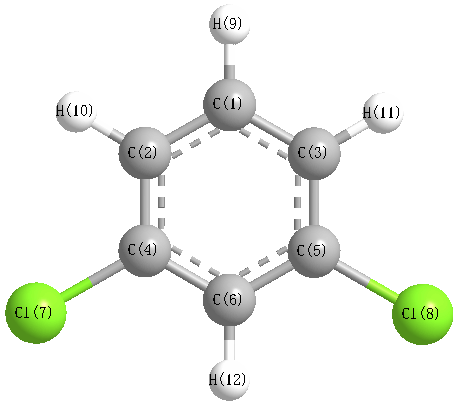
picture of 1,3-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 118.850 C1 C2 H10 121.056
C1 C3 C5 118.850 C1 C3 H11 121.056
C2 C1 C3 120.994 C2 C1 H9 119.503
C2 C4 C6 121.387 C2 C4 Cl7 119.645
C3 C1 H9 119.503 C3 C5 C6 121.387
C3 C5 Cl8 119.645 C4 C2 H10 120.094
C4 C6 C5 118.530 C4 C6 H12 120.735
C5 C3 H11 120.094 C5 C6 H12 120.735
C6 C4 Cl7 118.968 C6 C5 Cl8 118.968
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability