return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N (Cyclopropanecarbonitrile)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-209.536625
Energy at 298.15K-209.541912
HF Energy-208.802930
Nuclear repulsion energy147.436677
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3349 3145 2.69      
2 A' 3271 3072 0.43      
3 A' 3242 3045 1.93      
4 A' 2208 2073 6.54      
5 A' 1560 1465 0.76      
6 A' 1419 1332 0.31      
7 A' 1254 1178 1.18      
8 A' 1164 1094 0.36      
9 A' 1116 1048 8.11      
10 A' 994 933 26.58      
11 A' 839 788 1.52      
12 A' 778 730 3.96      
13 A' 520 488 0.71      
14 A' 205 193 3.51      
15 A" 3337 3134 0.25      
16 A" 3237 3040 5.82      
17 A" 1519 1427 2.48      
18 A" 1237 1162 0.76      
19 A" 1153 1083 3.25      
20 A" 1125 1056 6.52      
21 A" 940 883 1.26      
22 A" 862 809 10.54      
23 A" 537 504 0.22      
24 A" 213 200 2.06      

Unscaled Zero Point Vibrational Energy (zpe) 18038.0 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16941.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.53164 0.11429 0.10848

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -1.777 1.033 1.263
H2 -1.777 1.033 -1.263
H3 -1.251 -0.713 -1.254
H4 -1.251 -0.713 1.254
H5 0.274 1.673 0.000
C6 0.000 0.627 0.000
N7 2.025 -1.039 0.000
C8 1.117 -0.281 0.000
C9 -1.257 0.240 0.748
C10 -1.257 0.240 -0.748

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 C6 N7 C8 C9 C10
H12.52633.10841.82422.49212.21784.51023.42041.07942.2234
H22.52631.82423.10842.49212.21784.51023.42042.22341.0794
H33.10841.82422.50753.09692.22153.52282.71442.21711.0794
H41.82423.10842.50753.09692.22153.52282.71441.07942.2171
H52.49212.49213.09693.09691.08053.22722.12782.22622.2262
C62.21782.21782.22152.22151.08052.62181.43971.51341.5134
N74.51024.51023.52283.52283.22722.62181.18223.60083.6008
C83.42043.42042.71442.71442.12781.43971.18222.54342.5434
C91.07942.22342.21711.07942.22621.51343.60082.54341.4957
C102.22341.07941.07942.21712.22621.51343.60082.54341.4957

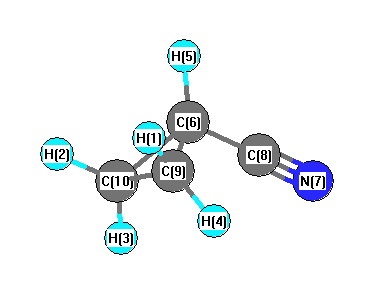
picture of Cyclopropanecarbonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C9 H4 115.351 H1 C9 C6 116.611
H1 C9 C10 118.517 H2 C10 H3 115.351
H2 C10 C6 116.611 H2 C10 C9 118.517
H3 C10 C6 116.934 H3 C10 C9 117.951
H4 C9 C6 116.934 H4 C9 C10 117.951
H5 C6 C8 114.442 H5 C6 C9 117.260
H5 C6 C10 117.260 C6 C8 N7 179.277
C6 C9 C10 60.387 C6 C10 C9 60.387
C8 C6 C9 118.895 C8 C6 C10 118.895
C9 C6 C10 59.226
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability