return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (1-Butene, 2-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-195.908988
Energy at 298.15K-195.919178
HF Energy-195.163255
Nuclear repulsion energy176.186766
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3322 3120 17.19      
2 A 3225 3029 15.27      
3 A 3222 3026 22.94      
4 A 3216 3021 23.04      
5 A 3213 3018 30.82      
6 A 3188 2994 18.60      
7 A 3166 2973 15.72      
8 A 3119 2929 28.19      
9 A 3107 2918 23.42      
10 A 3100 2911 29.25      
11 A 1730 1625 10.48      
12 A 1554 1460 1.23      
13 A 1548 1454 8.66      
14 A 1542 1448 11.81      
15 A 1531 1438 10.83      
16 A 1518 1426 3.37      
17 A 1488 1398 0.44      
18 A 1453 1364 4.60      
19 A 1451 1362 5.08      
20 A 1390 1305 1.50      
21 A 1341 1260 1.10      
22 A 1304 1225 1.15      
23 A 1142 1073 1.50      
24 A 1123 1055 4.76      
25 A 1077 1011 0.29      
26 A 1055 990 0.64      
27 A 1023 961 0.50      
28 A 971 912 1.15      
29 A 894 839 52.50      
30 A 816 766 1.43      
31 A 797 749 0.71      
32 A 731 687 0.23      
33 A 530 498 7.39      
34 A 433 407 1.52      
35 A 401 376 0.89      
36 A 272 256 0.16      
37 A 226 212 0.77      
38 A 181 170 0.43      
39 A 74 70 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 30735.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 28867.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.24236 0.12620 0.09537

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.936 1.348 0.037
H2 0.340 1.868 -0.713
H3 1.987 1.487 -0.211
H4 0.740 1.832 0.995
C5 -1.896 0.073 -0.389
H6 -1.899 1.161 -0.425
H7 -2.882 -0.250 -0.060
H8 -1.736 -0.299 -1.401
C9 -0.812 -0.455 0.552
H10 -0.901 -1.538 0.639
H11 -0.975 -0.039 1.551
C12 0.583 -0.112 0.099
C13 1.465 -1.068 -0.236
H14 1.203 -2.116 -0.187
H15 2.466 -0.824 -0.563

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15
C11.09021.08831.09163.13442.87824.14023.45242.56293.47412.80511.50342.48803.48132.7235
H21.09021.76341.75542.88502.36523.91073.07872.88463.86953.23932.15413.18004.11023.4336
H31.08831.76341.76804.13573.90505.17184.29753.49044.26833.76882.15082.60833.68802.3866
H41.09161.75541.76803.45803.07114.30924.05152.79863.76562.59832.14633.23274.14703.5299
C53.13442.88504.13573.45801.08851.08921.08951.52962.15502.15072.53293.55233.79914.4568
H62.87822.36523.90503.07111.08851.75811.76382.17873.06842.48982.83794.03964.51844.7973
H74.14023.91075.17184.30921.08921.75811.76472.16902.46472.50563.47154.42714.49315.4031
H83.45243.07874.29754.05151.08951.76381.76472.16602.52863.05932.76813.49193.66214.3173
C92.56292.88463.49042.79861.52962.17872.16902.16601.09101.09421.50542.48592.71363.4821
H103.47413.86954.26833.76562.15503.06842.46472.52861.09101.75622.12792.56642.33293.6466
H112.80513.23933.76882.59832.15072.48982.50563.05931.09421.75622.13043.19443.47494.1142
C121.50342.15412.15082.14632.53292.83793.47152.76811.50542.12792.13041.34332.11712.1198
C132.48803.18002.60833.23273.55234.03964.42713.49192.48592.56643.19441.34331.08151.0816
H143.48134.11023.68804.14703.79914.51844.49313.66212.71362.33293.47492.11711.08151.8464
H152.72353.43362.38663.52994.45684.79735.40314.31733.48213.64664.11422.11981.08161.8464

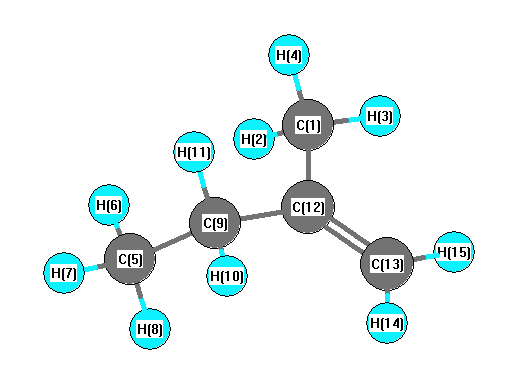
picture of 1-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 116.815 C1 C12 C13 121.753
H2 C1 H3 108.087 H2 C1 H4 107.138
H2 C1 C12 111.308 H3 C1 H4 108.401
H3 C1 C12 111.160 H4 C1 C12 110.598
C5 C9 H10 109.497 C5 C9 H11 108.972
C5 C9 C12 113.138 H6 C5 H7 107.667
H6 C5 H8 108.157 H6 C5 C9 111.536
H7 C5 H8 108.186 H7 C5 C9 110.713
H8 C5 C9 110.457 C9 C12 C13 121.431
H10 C9 H11 106.964 H10 C9 C12 109.037
H11 C9 C12 109.045 C12 C13 H14 121.263
C12 C13 H15 121.528 H14 C13 H15 117.209
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability