return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-306.566275
Energy at 298.15K-306.572608
HF Energy-305.557597
Nuclear repulsion energy268.467705
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3277 3078 0.00      
2 A1 3243 3046 5.16      
3 A1 3085 2898 5.90      
4 A1 1733 1628 144.63      
5 A1 1677 1575 61.99      
6 A1 1479 1389 18.67      
7 A1 1440 1353 1.19      
8 A1 1214 1140 6.46      
9 A1 973 914 3.85      
10 A1 907 852 10.24      
11 A1 789 741 2.54      
12 A1 504 473 1.80      
13 A2 1207 1133 0.00      
14 A2 977 917 0.00      
15 A2 740 695 0.00      
16 A2 336 315 0.00      
17 B1 3121 2931 3.12      
18 B1 993 933 12.05      
19 B1 950 892 15.47      
20 B1 764 717 56.11      
21 B1 544 511 14.41      
22 B1 203 190 2.30      
23 B1 97 91 1.34      
24 B2 3276 3077 12.59      
25 B2 3243 3046 12.65      
26 B2 1674 1573 3.11      
27 B2 1459 1370 38.24      
28 B2 1414 1328 1.38      
29 B2 1310 1230 24.20      
30 B2 1164 1094 7.16      
31 B2 1019 957 5.68      
32 B2 581 546 1.17      
33 B2 451 424 13.76      

Unscaled Zero Point Vibrational Energy (zpe) 22921.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 21527.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.17526 0.08858 0.05947

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.106
C2 0.000 0.000 -1.825
C3 0.000 1.254 0.334
C4 0.000 -1.254 0.334
C5 0.000 1.253 -1.015
C6 0.000 -1.253 -1.015
O7 0.000 0.000 2.347
H8 0.000 2.174 0.904
H9 0.000 -2.174 0.904
H10 0.000 2.193 -1.554
H11 0.000 -2.193 -1.554
H12 0.868 0.000 -2.494
H13 -0.868 0.000 -2.494

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.93131.47291.47292.46312.46311.24142.18302.18303.44753.44753.70333.7033
C22.93132.49682.49681.49271.49274.17263.48903.48902.21012.21011.09581.0958
C31.47292.49682.50821.34812.84682.37231.08203.47492.10833.93043.21293.2129
C41.47292.49682.50822.84681.34812.37233.47491.08203.93042.10833.21293.2129
C52.46311.49271.34812.84682.50663.58782.12773.92741.08383.48862.12462.1246
C62.46311.49272.84681.34812.50663.58783.92742.12773.48861.08382.12462.1246
O71.24144.17262.37232.37233.58783.58782.60932.60934.47554.47554.91874.9187
H82.18303.48901.08203.47492.12773.92742.60934.34742.45785.01114.12624.1262
H92.18303.48903.47491.08203.92742.12772.60934.34745.01112.45784.12624.1262
H103.44752.21012.10833.93041.08383.48864.47552.45785.01114.38672.53942.5394
H113.44752.21013.93042.10833.48861.08384.47555.01112.45784.38672.53942.5394
H123.70331.09583.21293.21292.12462.12464.91874.12624.12622.53942.53941.7358
H133.70331.09583.21293.21292.12462.12464.91874.12624.12622.53942.53941.7358

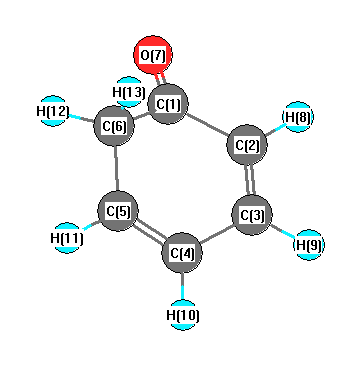
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 121.591 C1 C3 H8 116.569
C1 C4 C6 121.591 C1 C4 H9 116.569
C2 C5 C3 122.937 C2 C5 H10 117.253
C2 C6 C4 122.937 C2 C6 H11 117.253
C3 C1 C4 116.748 C3 C1 O7 121.626
C3 C5 H10 119.809 C4 C1 O7 121.626
C4 C6 H11 119.809 C5 C2 C6 114.196
C5 C2 H12 109.367 C5 C2 H13 109.367
C5 C3 H8 121.840 C6 C2 H12 109.367
C6 C2 H13 109.367 C6 C4 H9 121.840
H12 C2 H13 104.752
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability