return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,4-Pentadiene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-194.685473
Energy at 298.15K-194.693029
HF Energy-193.968646
Nuclear repulsion energy155.479672
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3332 3129 12.23      
2 A 3237 3040 2.51      
3 A 3225 3029 5.30      
4 A 3101 2913 19.48      
5 A 1723 1618 0.72      
6 A 1518 1425 6.20      
7 A 1486 1396 0.05      
8 A 1341 1260 0.36      
9 A 1286 1208 0.03      
10 A 1102 1035 3.73      
11 A 1029 967 21.15      
12 A 938 881 6.01      
13 A 926 870 42.76      
14 A 684 643 12.70      
15 A 376 353 0.50      
16 A 305 287 0.13      
17 A 90 85 0.05      
18 A 3332 3129 15.35      
19 A 3236 3039 7.65      
20 A 3225 3029 21.98      
21 A 3161 2969 11.66      
22 A 1714 1609 17.63      
23 A 1492 1401 2.66      
24 A 1351 1269 0.51      
25 A 1327 1246 3.60      
26 A 1201 1128 2.79      
27 A 1034 971 25.52      
28 A 997 936 2.85      
29 A 927 870 42.52      
30 A 914 858 8.14      
31 A 613 575 13.42      
32 A 458 430 2.51      
33 A 100 94 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 25389.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23846.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.65366 0.07907 0.07849

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.676
C2 0.000 1.240 -0.172
C3 0.000 -1.240 -0.172
C4 0.986 2.147 -0.186
C5 -0.986 -2.147 -0.186
H6 -0.881 -0.011 1.323
H7 0.881 0.011 1.323
H8 -0.858 1.375 -0.823
H9 0.858 -1.375 -0.823
H10 0.944 3.017 -0.826
H11 -0.944 -3.017 -0.826
H12 1.857 2.039 0.447
H13 -1.857 -2.039 0.447

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.50211.50212.51502.51501.09301.09302.20752.20753.49933.49932.76712.7671
C21.50212.47941.34013.52752.13912.12621.08542.82802.11544.40842.11433.8183
C31.50212.47943.52751.34012.12622.13912.82801.08544.40842.11543.81832.1143
C42.51501.34013.52754.72553.22792.61702.09853.58171.08035.54971.08215.0994
C52.51503.52751.34014.72552.61703.22793.58172.09855.54971.08035.09941.0821
H61.09302.13912.12623.22792.61701.76212.55433.08034.13673.69483.53062.4145
H71.09302.12622.13912.61703.22791.76213.08032.55433.69484.13672.41453.5306
H82.20751.08542.82802.09853.58172.55433.08033.24232.43774.39273.07013.7766
H92.20752.82801.08543.58172.09853.08032.55433.24234.39272.43773.77663.0701
H103.49932.11544.40841.08035.54974.13673.69482.43774.39276.32181.84675.9177
H113.49934.40842.11545.54971.08033.69484.13674.39272.43776.32185.91771.8467
H122.76712.11433.81831.08215.09943.53062.41453.07013.77661.84675.91775.5152
H132.76713.81832.11435.09941.08212.41453.53063.77663.07015.91771.84675.5152

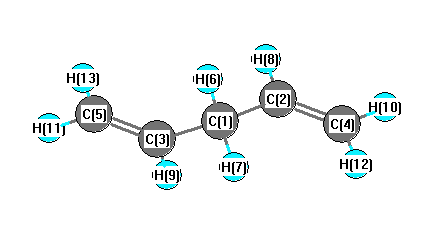
picture of 1,4-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.369 C1 C2 H8 116.186
C1 C3 C5 124.369 C1 C3 H9 116.186
C2 C1 C3 111.240 C2 C1 H6 110.035
C2 C1 H7 109.010 C2 C4 H10 121.477
C2 C4 H12 121.227 C3 C1 H6 109.010
C3 C1 H7 110.035 C3 C5 H11 121.477
C3 C5 H13 121.227 C4 C2 H8 119.433
C5 C3 H9 119.433 H6 C1 H7 107.433
H10 C4 H12 117.296 H11 C5 H13 117.296
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31+G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.220      
2 C -0.074      
3 C -0.074      
4 C -0.387      
5 C -0.387      
6 H 0.140      
7 H 0.140      
8 H 0.154      
9 H 0.154      
10 H 0.143      
11 H 0.143      
12 H 0.134      
13 H 0.134      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.859 2.469 0.000
y 2.469 10.633 0.000
z 0.000 0.000 6.947


<r2> (average value of r2) Å2
<r2> 157.913
(<r2>)1/2 12.566