return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-235.092792
Energy at 298.15K-235.105269
HF Energy-234.198785
Nuclear repulsion energy228.660072
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3330 3127 15.61      
2 A 3231 3035 4.59      
3 A 3218 3022 23.37      
4 A 3210 3015 34.39      
5 A 3208 3013 44.98      
6 A 3171 2978 52.48      
7 A 3155 2963 0.37      
8 A 3142 2951 1.03      
9 A 3113 2924 35.80      
10 A 3108 2919 64.35      
11 A 3098 2910 13.32      
12 A 3093 2905 4.11      
13 A 1721 1617 9.77      
14 A 1553 1459 6.89      
15 A 1546 1452 7.94      
16 A 1542 1448 1.90      
17 A 1532 1439 0.76      
18 A 1525 1433 2.17      
19 A 1492 1401 0.84      
20 A 1457 1368 3.33      
21 A 1436 1348 0.50      
22 A 1384 1299 0.36      
23 A 1357 1275 0.78      
24 A 1344 1263 0.17      
25 A 1338 1257 0.56      
26 A 1295 1216 1.56      
27 A 1278 1200 0.15      
28 A 1228 1153 0.24      
29 A 1156 1085 2.70      
30 A 1110 1043 0.21      
31 A 1072 1007 3.75      
32 A 1061 997 2.35      
33 A 1031 968 22.17      
34 A 968 909 0.77      
35 A 947 890 0.89      
36 A 929 873 1.01      
37 A 926 870 45.35      
38 A 817 767 0.89      
39 A 749 703 1.74      
40 A 656 616 13.07      
41 A 464 436 0.87      
42 A 365 343 0.08      
43 A 364 341 0.69      
44 A 254 239 0.00      
45 A 179 168 0.05      
46 A 131 123 0.08      
47 A 93 88 0.02      
48 A 77 72 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 37226.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 34962.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.44167 0.04205 0.04130

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.050 -0.221 -0.449
C2 2.028 -0.168 0.416
C3 0.770 0.614 0.198
C4 -0.466 -0.283 0.118
C5 -1.762 0.497 -0.068
C6 -2.986 -0.406 -0.149
H7 3.028 0.336 -1.377
H8 3.930 -0.816 -0.250
H9 2.088 -0.743 1.336
H10 0.634 1.328 1.016
H11 0.860 1.196 -0.722
H12 -0.341 -0.986 -0.710
H13 -0.536 -0.885 1.030
H14 -1.878 1.201 0.760
H15 -1.689 1.098 -0.978
H16 -3.900 0.171 -0.284
H17 -2.901 -1.100 -0.985
H18 -3.093 -0.995 0.763

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34052.51263.56234.88006.04671.08221.08042.09443.22232.62313.48593.93515.26924.94716.96306.03906.3089
C21.34051.49702.51443.87805.05142.11432.11681.08622.13102.12682.74722.73144.15264.16655.97855.20765.1983
C32.51261.49701.52982.54863.90802.76673.49682.20731.09431.09282.14912.15472.76962.76834.71574.22024.2224
C43.56232.51441.52981.52412.53713.85044.44332.86672.14772.15731.09331.09412.14642.14593.48692.79452.7964
C54.88003.87802.54861.52411.52404.96775.84394.28132.75772.79082.15222.14911.09291.09282.17362.16522.1652
C66.04675.05143.90802.53711.52406.18286.92895.29764.17964.20592.76592.76082.15352.15311.08921.08981.0899
H71.08222.11432.76673.85044.96776.18281.84643.06743.52742.42273.67924.46975.42014.79407.01526.11206.6187
H81.08042.11683.49684.44335.84396.92891.84642.43214.13043.70084.29864.64576.22985.97987.89176.87567.0974
H92.09441.08622.20732.86674.28135.29763.06742.43212.55003.08323.18522.64524.45334.79656.26995.51365.2183
H103.22232.13101.09432.14772.75774.17963.52744.13042.55001.75823.04692.50262.52763.06994.85624.73174.3984
H112.62312.12681.09282.15732.79084.20592.42273.70083.08321.75822.49063.05733.11302.56294.88854.41364.7569
H123.48592.74722.14911.09332.15222.76593.67924.29863.18523.04692.49061.75353.05052.49633.76672.57703.1214
H133.93512.73142.15471.09412.14912.76084.46974.64572.64522.50263.05731.75352.49443.04833.76263.11402.5733
H145.26924.15262.76962.14641.09292.15355.42016.22984.45332.52763.11303.05052.49441.75182.49793.06342.5092
H154.94714.16652.76832.14591.09282.15314.79405.97984.79653.06992.56292.49633.04831.75182.49642.50993.0631
H166.96305.97854.71573.48692.17361.08927.01527.89176.26994.85624.88853.76673.76262.49792.49641.76231.7622
H176.03905.20764.22022.79452.16521.08986.11206.87565.51364.73174.41362.57703.11403.06342.50991.76231.7613
H186.30895.19834.22242.79642.16521.08996.61877.09745.21834.39844.75693.12142.57332.50923.06311.76221.7613

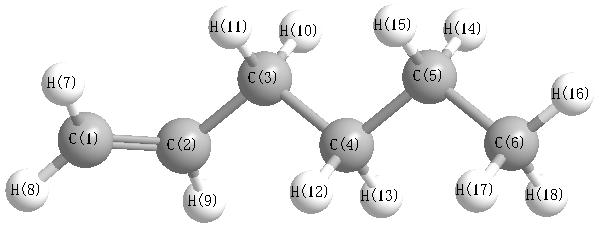
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.535 C1 C2 H9 118.952
C2 C1 H7 121.180 C2 C1 H8 121.566
C2 C3 C4 112.335 C2 C3 H10 109.661
C2 C3 H11 109.420 C3 C2 H9 116.500
C3 C4 C5 113.132 C3 C4 H12 108.889
C3 C4 H13 109.281 C4 C3 H10 108.723
C4 C3 H11 109.559 C4 C5 C6 112.683
C4 C5 H14 109.087 C4 C5 H15 109.052
C5 C4 H12 109.518 C5 C4 H13 109.234
C5 C6 H16 111.478 C5 C6 H17 110.764
C5 C6 H18 110.767 C6 C5 H14 109.658
C6 C5 H15 109.631 H7 C1 H8 117.254
H10 C3 H11 106.996 H12 C4 H13 106.575
H14 C5 H15 106.540 H16 C6 H17 107.945
H16 C6 H18 107.937 H17 C6 H18 107.808
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability