return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,2-Butadiene, 3-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-194.675528
Energy at 298.15K-194.682509
HF Energy-193.955305
Nuclear repulsion energy157.174083
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3230 3033 18.32      
2 A1 3215 3019 5.91      
3 A1 3105 2916 50.57      
4 A1 2063 1937 15.39      
5 A1 1557 1462 0.16      
6 A1 1517 1425 2.55      
7 A1 1454 1366 0.74      
8 A1 1341 1259 13.53      
9 A1 1050 986 5.29      
10 A1 750 704 0.02      
11 A1 344 323 0.05      
12 A2 3184 2990 0.00      
13 A2 1517 1425 0.00      
14 A2 1025 963 0.00      
15 A2 629 590 0.00      
16 A2 150 141 0.00      
17 B1 3306 3105 2.56      
18 B1 3184 2990 37.40      
19 B1 1532 1439 16.53      
20 B1 1113 1046 0.85      
21 B1 1040 977 0.35      
22 B1 394 370 0.76      
23 B1 246 231 7.67      
24 B1 157 147 1.09      
25 B2 3229 3033 14.14      
26 B2 3103 2915 21.74      
27 B2 1533 1440 7.07      
28 B2 1438 1351 5.29      
29 B2 1257 1180 4.47      
30 B2 996 936 0.14      
31 B2 877 824 59.16      
32 B2 597 561 11.85      
33 B2 178 168 1.02      

Unscaled Zero Point Vibrational Energy (zpe) 25155.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23625.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.27738 0.11985 0.08792

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.250
C2 0.000 0.000 0.934
H3 -0.924 0.000 2.812
H4 0.924 0.000 2.812
C5 0.000 0.000 -0.383
C6 0.000 1.283 -1.175
C7 0.000 -1.283 -1.175
H8 0.000 2.153 -0.523
H9 0.000 -2.153 -0.523
H10 0.880 1.328 -1.819
H11 -0.880 1.328 -1.819
H12 -0.880 -1.328 -1.819
H13 0.880 -1.328 -1.819

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 H8 H9 H10 H11 H12 H13
C11.31591.08201.08202.63253.65693.65693.51023.51024.36974.36974.36974.3697
C21.31592.09332.09331.31662.46822.46822.59932.59933.18083.18083.18083.1808
H31.08202.09331.84903.32584.28894.28894.07564.07565.14464.81814.81815.1446
H41.08202.09331.84903.32584.28894.28894.07564.07564.81815.14465.14464.8181
C52.63251.31663.32583.32581.50751.50752.15722.15722.14502.14502.14502.1450
C63.65692.46824.28894.28891.50752.56521.08713.49651.09131.09132.82932.8293
C73.65692.46824.28894.28891.50752.56523.49651.08712.82932.82931.09131.0913
H83.51022.59934.07564.07562.15721.08713.49654.30521.77021.77023.81703.8170
H93.51022.59934.07564.07562.15723.49651.08714.30523.81703.81701.77021.7702
H104.36973.18085.14464.81812.14501.09132.82931.77023.81701.75913.18592.6563
H114.36973.18084.81815.14462.14501.09132.82931.77023.81701.75912.65633.1859
H124.36973.18084.81815.14462.14502.82931.09133.81701.77023.18592.65631.7591
H134.36973.18085.14464.81812.14502.82931.09133.81701.77022.65633.18591.7591

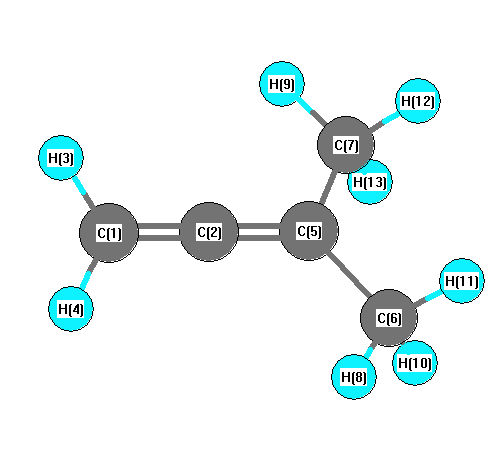
picture of 1,2-Butadiene, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C5 180.000 C2 C1 H3 121.305
C2 C1 H4 121.305 C2 C5 C6 121.698
C2 C5 C7 121.698 H3 C1 H4 117.389
C5 C6 H8 111.458 C5 C6 H10 110.228
C5 C6 H11 110.228 C5 C7 H9 111.458
C5 C7 H12 110.228 C5 C7 H13 110.228
C6 C5 C7 116.604 H8 C6 H10 108.705
H8 C6 H11 108.705 H9 C7 H12 108.705
H9 C7 H13 108.705 H10 C6 H11 107.407
H12 C7 H13 107.407
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability