return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSCH3 (Disulfide, dimethyl)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-874.841747
Energy at 298.15K-874.847992
HF Energy-874.258926
Nuclear repulsion energy222.459063
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3248 3051 3.46      
2 A 3236 3039 5.98      
3 A 3130 2940 20.22      
4 A 1515 1423 0.64      
5 A 1497 1406 6.97      
6 A 1423 1337 1.28      
7 A 1017 955 2.90      
8 A 1012 950 2.13      
9 A 748 703 0.01      
10 A 525 494 0.29      
11 A 243 228 1.42      
12 A 159 149 0.00      
13 A 99 93 1.23      
14 B 3248 3050 6.73      
15 B 3236 3039 4.94      
16 B 3128 2938 19.92      
17 B 1519 1427 19.40      
18 B 1489 1399 11.68      
19 B 1417 1330 2.56      
20 B 1019 957 13.76      
21 B 1008 947 1.74      
22 B 744 699 1.82      
23 B 272 255 0.94      
24 B 174 164 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 17551.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16484.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.26777 0.09438 0.08582

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 1.028 -0.506
S2 0.000 -1.028 -0.506
C3 1.182 1.400 0.813
C4 -1.182 -1.400 0.813
H5 1.277 2.484 0.849
H6 0.826 1.050 1.779
H7 2.152 0.963 0.597
H8 -1.277 -2.484 0.849
H9 -0.826 -1.050 1.779
H10 -2.152 -0.963 0.597

Atom - Atom Distances (Å)
  S1 S2 C3 C4 H5 H6 H7 H8 H9 H10
S12.05561.80983.00522.36392.42982.41973.97493.19713.1329
S22.05563.00521.80983.97493.19713.13292.36392.42982.4197
C31.80983.00523.66391.08861.08681.08624.59673.31144.0922
C43.00521.80983.66394.59673.31144.09221.08861.08681.0862
H52.36393.97491.08864.59671.76721.77255.58584.21614.8690
H62.42983.19711.08683.31141.76721.77834.21612.67213.7842
H72.41973.13291.08624.09221.77251.77834.86903.78424.7162
H83.97492.36394.59671.08865.58584.21614.86901.76721.7725
H93.19712.42983.31141.08684.21612.67213.78421.76721.7783
H103.13292.41974.09221.08624.86903.78424.71621.77251.7783

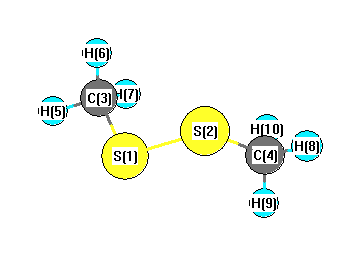
picture of Disulfide, dimethyl state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 101.866 S1 C3 H5 106.627
S1 C3 H6 111.590 S1 C3 H7 110.853
S2 S1 C3 101.866 S2 C4 H8 106.627
S2 C4 H9 111.590 S2 C4 H10 110.853
H5 C3 H6 108.654 H5 C3 H7 109.172
H6 C3 H7 109.844 H8 C4 H9 108.654
H8 C4 H10 109.172 H9 C4 H10 109.844
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability