return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-531.229238
Energy at 298.15K 
HF Energy-530.625216
Nuclear repulsion energy155.230693
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3799 3568 53.00      
2 A 3650 3428 81.07      
3 A 3268 3069 1.72      
4 A 3204 3009 10.31      
5 A 3119 2929 17.37      
6 A 1685 1582 173.69      
7 A 1528 1435 11.74      
8 A 1527 1434 12.19      
9 A 1446 1358 22.66      
10 A 1424 1337 360.30      
11 A 1360 1277 19.23      
12 A 1060 996 9.78      
13 A 1040 977 32.37      
14 A 1024 962 2.85      
15 A 765 718 3.09      
16 A 587 551 3.18      
17 A 483 454 35.70      
18 A 440 413 1.07      
19 A 388 364 0.94      
20 A 74 69 0.66      
21 A 130i 122i 183.70      

Unscaled Zero Point Vibrational Energy (zpe) 15869.1 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 14904.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.32556 0.16582 0.11216

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.270 0.055 -0.013
S2 -1.363 -0.109 0.001
C3 1.226 -1.110 0.001
N4 0.888 1.261 -0.003
H5 0.717 -2.012 -0.318
H6 2.079 -0.927 -0.652
H7 1.597 -1.263 1.015
H8 1.891 1.345 -0.002
H9 0.335 2.102 0.038

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.64211.50731.35512.13692.15512.13452.07122.0483
S21.64212.77632.63592.83783.59813.33573.56452.7880
C31.50732.77632.39541.08351.09031.09042.54363.3337
N41.35512.63592.39543.29282.57442.81231.00631.0074
H52.13692.83781.08353.29281.77291.76483.57014.1470
H62.15513.59811.09032.57441.77291.76782.37043.5628
H72.13453.33571.09042.81231.76481.76782.81423.7241
H82.07123.56452.54361.00633.57012.37042.81421.7313
H92.04832.78803.33371.00744.14703.56283.72411.7313

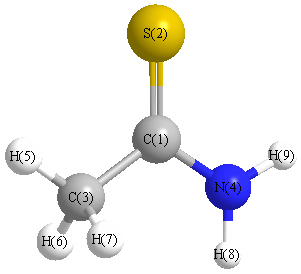
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.061 C1 C3 H6 111.108
C1 C3 H7 109.462 C1 N4 H8 121.894
C1 N4 H9 119.495 S2 C1 C3 123.598
S2 C1 N4 122.866 C3 C1 N4 113.513
H5 C3 H6 109.287 H5 C3 H7 108.543
H6 C3 H7 108.324 H8 N4 H9 118.574
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability