return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (E)-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-195.906327
Energy at 298.15K-195.916449
HF Energy-195.162597
Nuclear repulsion energy171.063410
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3244 3047 22.99      
2 A' 3216 3021 29.78      
3 A' 3211 3016 20.00      
4 A' 3204 3009 19.89      
5 A' 3121 2931 22.26      
6 A' 3108 2919 34.36      
7 A' 3099 2911 30.65      
8 A' 1751 1645 0.07      
9 A' 1555 1461 5.75      
10 A' 1544 1450 10.68      
11 A' 1527 1434 2.72      
12 A' 1463 1374 8.27      
13 A' 1455 1367 0.19      
14 A' 1414 1328 6.36      
15 A' 1359 1276 0.81      
16 A' 1352 1270 0.13      
17 A' 1179 1107 0.67      
18 A' 1128 1059 4.23      
19 A' 1061 996 2.03      
20 A' 980 920 5.95      
21 A' 887 833 0.04      
22 A' 589 553 0.51      
23 A' 372 349 0.19      
24 A' 213 200 0.25      
25 A" 3217 3021 29.46      
26 A" 3188 2995 17.74      
27 A" 3141 2950 14.81      
28 A" 1543 1449 7.63      
29 A" 1521 1429 6.48      
30 A" 1315 1235 0.56      
31 A" 1132 1063 2.51      
32 A" 1081 1015 0.87      
33 A" 1009 947 41.17      
34 A" 837 786 1.74      
35 A" 709 666 0.17      
36 A" 308 289 1.95      
37 A" 217 204 1.67      
38 A" 190 179 2.16      
39 A" 109 103 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 30772.5 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 28901.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.44683 0.08256 0.07247

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.713 -1.238 0.000
C2 -0.974 -0.186 0.000
H3 -0.289 1.785 0.000
C4 0.000 0.737 0.000
H5 -2.932 -0.280 0.877
H6 -2.932 -0.280 -0.877
H7 -2.599 1.215 0.000
C8 -2.437 0.138 0.000
H9 1.919 0.967 -0.870
H10 1.919 0.967 0.870
C11 1.479 0.472 0.000
H12 2.963 -1.095 0.000
H13 1.496 -1.509 0.882
H14 1.496 -1.509 -0.882
C15 1.879 -0.997 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 H7 C8 H9 H10 C11 H12 H13 H14 C15
H11.08423.05262.09992.57092.57093.09362.20573.54223.54222.78033.67932.39382.39382.6035
C21.08422.08631.34192.14742.14742.14461.49843.23343.23342.53974.04112.93742.93742.9663
H33.05262.08631.08723.46693.46692.37862.70692.51032.51032.20234.34423.84863.84863.5270
C42.09991.34191.08723.22523.22522.64212.51002.11932.11931.50233.48362.83852.83852.5566
H52.57092.14743.46693.22521.75471.76531.09055.30465.00844.55976.01564.59504.92014.9425
H62.57092.14743.46693.22521.75471.76531.09055.00845.30464.55976.01564.92014.59504.9425
H73.09362.14462.37862.64211.76531.76531.08904.60694.60694.14446.02224.99574.99574.9939
C82.20571.49842.70692.51001.09051.09051.08904.51884.51883.93045.53944.35404.35404.4629
H93.54223.23342.51032.11935.30465.00844.60694.51881.74091.09362.46983.06222.51152.1484
H103.54223.23342.51032.11935.00845.30464.60694.51881.74091.09362.46982.51153.06222.1484
C112.78032.53972.20231.50234.55974.55974.14443.93041.09361.09362.15842.16812.16811.5224
H123.67934.04114.34423.48366.01566.01566.02225.53942.46982.46982.15841.76121.76121.0887
H132.39382.93743.84862.83854.59504.92014.99574.35403.06222.51152.16811.76121.76311.0890
H142.39382.93743.84862.83854.92014.59504.99574.35402.51153.06222.16811.76121.76311.0890
C152.60352.96633.52702.55664.94254.94254.99394.46292.14842.14841.52241.08871.08901.0890

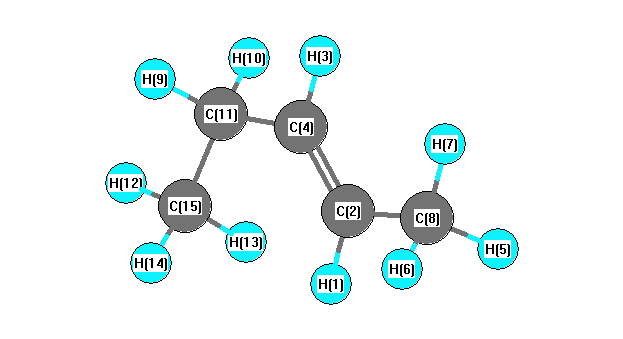
picture of 2-Pentene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 119.511 H1 C2 C8 116.395
C2 C4 H3 118.005 C2 C4 C11 126.405
C2 C8 H5 111.110 C2 C8 H6 111.110
C2 C8 H7 110.979 H3 C4 C11 115.590
C4 C2 C8 124.095 C4 C11 H9 108.424
C4 C11 H10 108.424 C4 C11 C15 115.396
H5 C8 H6 107.127 H5 C8 H7 108.181
H6 C8 H7 108.181 H9 C11 H10 105.483
H9 C11 C15 109.326 H10 C11 C15 109.326
C11 C15 H12 110.407 C11 C15 H13 111.160
C11 C15 H14 111.160 H12 C15 H13 107.946
H12 C15 H14 107.946 H13 C15 H14 108.093
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability