return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5COOH (benzoic acid)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-419.674823
Energy at 298.15K-419.681396
HF Energy-418.348646
Nuclear repulsion energy404.319095
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3793 3563 102.97      
2 A 3292 3092 2.79      
3 A 3284 3084 6.35      
4 A 3275 3075 9.84      
5 A 3265 3066 9.37      
6 A 3254 3056 0.16      
7 A 1798 1689 301.81      
8 A 1676 1574 7.57      
9 A 1655 1554 6.90      
10 A 1550 1456 5.55      
11 A 1505 1413 13.80      
12 A 1477 1387 1.97      
13 A 1390 1306 161.00      
14 A 1363 1280 2.33      
15 A 1229 1154 41.19      
16 A 1213 1139 120.01      
17 A 1207 1134 1.88      
18 A 1131 1062 35.82      
19 A 1105 1038 125.38      
20 A 1059 995 22.21      
21 A 1020 958 1.48      
22 A 923 867 0.11      
23 A 912 856 0.30      
24 A 889 835 0.50      
25 A 845 794 0.21      
26 A 780 733 9.33      
27 A 760 713 5.53      
28 A 708 665 150.55      
29 A 632 594 48.10      
30 A 628 590 0.17      
31 A 574 539 79.60      
32 A 499 468 5.94      
33 A 409 384 3.39      
34 A 385 362 5.44      
35 A 373 350 0.80      
36 A 350 328 3.49      
37 A 218 205 1.35      
38 A 133 125 1.01      
39 A 31 29 0.58      

Unscaled Zero Point Vibrational Energy (zpe) 25293.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23755.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.12848 0.04089 0.03105

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.219 0.030 -0.005
C2 -0.509 1.226 0.031
C3 -1.902 1.185 0.034
C4 -2.569 -0.043 0.004
C5 -1.839 -1.233 -0.033
C6 -0.444 -1.204 -0.039
C7 1.700 0.122 -0.008
O8 2.338 1.162 -0.079
O9 2.303 -1.096 0.083
H10 0.026 2.165 0.054
H11 -2.467 2.108 0.063
H12 -3.651 -0.072 0.009
H13 -2.355 -2.184 -0.058
H14 0.126 -2.121 -0.070
H15 3.259 -0.922 0.055

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 O9 H10 H11 H12 H13 H14 H15
C11.40032.41552.78922.41431.40091.48372.40412.37092.14453.39613.87143.39502.15393.1864
C21.40031.39342.41942.79562.43112.46972.85063.64751.08142.14733.39933.87763.40774.3375
C32.41551.39341.39792.42002.79973.75584.24224.78462.16271.08202.15403.40043.88015.5749
C42.78922.41941.39791.39722.42224.27235.05434.98583.40762.15381.08222.15263.40455.8946
C52.41432.79562.42001.39721.39493.78884.81514.14553.87683.40072.15311.08202.15645.1078
C61.40092.43112.79972.42221.39492.52053.65242.75183.40263.88173.40142.14741.08043.7148
C71.48372.46973.75584.27233.78882.52051.22281.36262.64224.61615.35444.66422.74021.8776
O82.40412.85064.24225.05434.81513.65241.22282.26472.52444.89946.11605.76363.95872.2824
O92.37093.64754.78464.98584.14552.75181.36262.26473.97815.74626.04234.78492.41060.9721
H102.14451.08142.16273.40763.87683.40262.64222.52443.97812.49314.30414.95894.28894.4705
H113.39612.14731.08202.15383.40073.88174.61614.89945.74622.49312.48124.29474.96216.4780
H123.87143.39932.15401.08222.15313.40145.35446.11606.04234.30412.48122.47914.29816.9625
H133.39503.87763.40042.15261.08202.14744.66425.76364.78494.95894.29472.47912.48175.7547
H142.15393.40773.88013.40452.15641.08042.74023.95872.41064.28894.96214.29812.48173.3565
H153.18644.33755.57495.89465.10783.71481.87762.28240.97214.47056.47806.96255.75473.3565

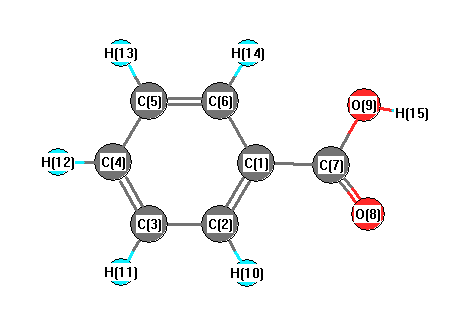
picture of benzoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.676 C1 C2 H10 119.014
C1 C6 C5 119.434 C1 C6 H14 119.908
C1 C7 O8 125.034 C1 C7 O9 112.744
C2 C1 C6 120.432 C2 C1 C7 117.792
C2 C3 C4 120.168 C2 C3 H11 119.797
C3 C2 H10 121.310 C3 C4 C5 119.946
C3 C4 H12 120.042 C4 C3 H11 120.035
C4 C5 C6 120.343 C4 C5 H13 119.975
C5 C4 H12 120.012 C5 C6 H14 120.658
C6 C1 C7 121.776 C6 C5 H13 119.682
C7 O9 H15 105.852 O8 C7 O9 122.222
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability