return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6O2S (Dimethyl sulfone)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-627.251441
Energy at 298.15K 
HF Energy-626.405781
Nuclear repulsion energy272.429666
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3268 3070 0.05      
2 A1 3144 2953 1.83      
3 A1 1506 1415 3.98      
4 A1 1410 1325 11.95      
5 A1 1162 1091 179.71      
6 A1 1047 983 0.42      
7 A1 709 666 8.65      
8 A1 476 447 27.72      
9 A1 274 258 2.16      
10 A2 3270 3071 0.00      
11 A2 1494 1403 0.00      
12 A2 976 916 0.00      
13 A2 297 279 0.00      
14 A2 207 194 0.00      
15 B1 3274 3075 0.56      
16 B1 1510 1418 11.18      
17 B1 1337 1256 281.94      
18 B1 1032 970 0.23      
19 B1 362 340 1.93      
20 B1 239 224 0.35      
21 B2 3267 3068 1.10      
22 B2 3142 2951 0.04      
23 B2 1497 1406 6.40      
24 B2 1392 1307 15.00      
25 B2 984 924 54.20      
26 B2 784 736 45.74      
27 B2 453 426 39.55      

Unscaled Zero Point Vibrational Energy (zpe) 19256.0 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 18085.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.14971 0.13924 0.13801

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.188
O2 -1.281 0.000 0.913
O3 1.281 0.000 0.913
C4 0.000 1.401 -0.913
C5 0.000 -1.401 -0.913
H6 0.000 2.282 -0.277
H7 0.000 -2.282 -0.277
H8 0.899 1.380 -1.523
H9 -0.899 1.380 -1.523
H10 -0.899 -1.380 -1.523
H11 0.899 -1.380 -1.523

Atom - Atom Distances (Å)
  S1 O2 O3 C4 C5 H6 H7 H8 H9 H10 H11
S11.47201.47201.78231.78232.32882.32882.37472.37472.37472.3747
O21.47202.56222.63442.63442.87472.87473.54812.82542.82543.5481
O31.47202.56222.63442.63442.87472.87472.82543.54813.54812.8254
C41.78232.63442.63442.80281.08673.73801.08611.08612.98612.9861
C51.78232.63442.63442.80283.73801.08672.98612.98611.08611.0861
H62.32882.87472.87471.08673.73804.56411.78141.78143.97163.9716
H72.32882.87472.87473.73801.08674.56413.97163.97161.78141.7814
H82.37473.54812.82541.08612.98611.78143.97161.79753.29432.7607
H92.37472.82543.54811.08612.98611.78143.97161.79752.76073.2943
H102.37472.82543.54812.98611.08613.97161.78143.29432.76071.7975
H112.37473.54812.82542.98611.08613.97161.78142.76073.29431.7975

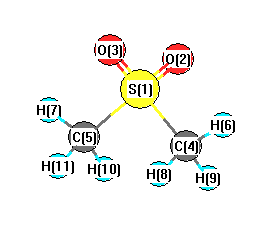
picture of Dimethyl sulfone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C4 H6 105.976 S1 C4 H8 109.355
S1 C4 H9 109.355 S1 C5 H7 105.976
S1 C5 H10 109.355 S1 C5 H11 109.355
O2 S1 O3 120.991 O2 S1 C4 107.715
O2 S1 C5 107.715 O3 S1 C4 107.715
O3 S1 C5 107.715 C4 S1 C5 103.681
H6 C4 H8 110.146 H6 C4 H9 110.146
H7 C5 H10 110.146 H7 C5 H11 110.146
H8 C4 H9 111.694 H10 C5 H11 111.694
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability