Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -498.733458 |
| Energy at 298.15K | |
| HF Energy | -498.466236 |
| Nuclear repulsion energy | 45.507551 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3293 |
3093 |
7.98 |
|
|
|
| 2 |
A' |
1502 |
1411 |
16.96 |
|
|
|
| 3 |
A' |
884 |
830 |
40.33 |
|
|
|
| 4 |
A' |
192 |
180 |
92.67 |
|
|
|
| 5 |
A" |
3449 |
3240 |
0.05 |
|
|
|
| 6 |
A" |
1053 |
989 |
0.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5186.8 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4871.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.006 |
1.114 |
0.000 |
| Cl2 |
-0.006 |
-0.583 |
0.000 |
| H3 |
0.073 |
1.615 |
0.946 |
| H4 |
0.073 |
1.615 |
-0.946 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.6975 | 1.0732 | 1.0732 |
Cl2 | 1.6975 | | 2.3947 | 2.3947 | H3 | 1.0732 | 2.3947 | | 1.8912 | H4 | 1.0732 | 2.3947 | 1.8912 | |
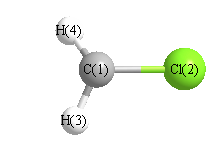 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.841 |
|
Br2 |
C1 |
H4 |
117.841 |
| H3 |
C1 |
H4 |
123.565 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -498.733452 |
| Energy at 298.15K | |
| HF Energy | -498.466180 |
| Nuclear repulsion energy | 45.519807 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3297 |
3096 |
7.44 |
|
|
|
| 2 |
A1 |
1502 |
1411 |
17.18 |
|
|
|
| 3 |
A1 |
885 |
832 |
39.75 |
|
|
|
| 4 |
B1 |
138i |
129i |
99.17 |
|
|
|
| 5 |
B2 |
3454 |
3244 |
0.11 |
|
|
|
| 6 |
B2 |
1051 |
987 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5025.9 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 4720.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.113 |
| Cl2 |
0.000 |
0.000 |
0.583 |
| H3 |
0.000 |
0.947 |
-1.618 |
| H4 |
0.000 |
-0.947 |
-1.618 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.6967 | 1.0728 | 1.0728 |
Cl2 | 1.6967 | | 2.3961 | 2.3961 | H3 | 1.0728 | 2.3961 | | 1.8937 | H4 | 1.0728 | 2.3961 | 1.8937 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
118.042 |
|
Br2 |
C1 |
H4 |
118.042 |
| H3 |
C1 |
H4 |
123.916 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
