Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -477.197603 |
| Energy at 298.15K | -477.203830 |
| HF Energy | -476.748423 |
| Nuclear repulsion energy | 107.810714 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3225 |
3029 |
19.17 |
|
|
|
| 2 |
A' |
3151 |
2959 |
23.17 |
|
|
|
| 3 |
A' |
3127 |
2937 |
20.21 |
|
|
|
| 4 |
A' |
2831 |
2659 |
10.68 |
|
|
|
| 5 |
A' |
1549 |
1455 |
2.65 |
|
|
|
| 6 |
A' |
1535 |
1442 |
3.93 |
|
|
|
| 7 |
A' |
1463 |
1374 |
5.14 |
|
|
|
| 8 |
A' |
1365 |
1282 |
41.70 |
|
|
|
| 9 |
A' |
1151 |
1081 |
2.94 |
|
|
|
| 10 |
A' |
1035 |
972 |
1.73 |
|
|
|
| 11 |
A' |
901 |
846 |
3.12 |
|
|
|
| 12 |
A' |
717 |
673 |
0.42 |
|
|
|
| 13 |
A' |
315 |
295 |
2.39 |
|
|
|
| 14 |
A" |
3233 |
3036 |
22.91 |
|
|
|
| 15 |
A" |
3209 |
3014 |
0.49 |
|
|
|
| 16 |
A" |
1535 |
1442 |
9.32 |
|
|
|
| 17 |
A" |
1303 |
1224 |
0.61 |
|
|
|
| 18 |
A" |
1086 |
1020 |
1.11 |
|
|
|
| 19 |
A" |
814 |
764 |
2.52 |
|
|
|
| 20 |
A" |
265 |
249 |
0.86 |
|
|
|
| 21 |
A" |
120 |
113 |
20.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16964.2 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 15932.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.514 |
0.681 |
0.000 |
| C2 |
0.000 |
0.825 |
0.000 |
| S3 |
-0.757 |
-0.832 |
0.000 |
| H4 |
1.981 |
1.665 |
0.000 |
| H5 |
1.854 |
0.143 |
0.882 |
| H6 |
1.854 |
0.143 |
-0.882 |
| H7 |
-0.322 |
1.372 |
0.883 |
| H8 |
-0.322 |
1.372 |
-0.883 |
| H9 |
-2.024 |
-0.425 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5206 | 2.7286 | 1.0885 | 1.0883 | 1.0883 | 2.1512 | 2.1512 | 3.7066 |
C2 | 1.5206 | | 1.8214 | 2.1517 | 2.1638 | 2.1638 | 1.0879 | 1.0879 | 2.3785 | S3 | 2.7286 | 1.8214 | | 3.7051 | 2.9234 | 2.9234 | 2.4142 | 2.4142 | 1.3310 | H4 | 1.0885 | 2.1517 | 3.7051 | | 1.7633 | 1.7633 | 2.4839 | 2.4839 | 4.5172 | H5 | 1.0883 | 2.1638 | 2.9234 | 1.7633 | | 1.7649 | 2.4994 | 3.0602 | 4.0176 | H6 | 1.0883 | 2.1638 | 2.9234 | 1.7633 | 1.7649 | | 3.0602 | 2.4994 | 4.0176 | H7 | 2.1512 | 1.0879 | 2.4142 | 2.4839 | 2.4994 | 3.0602 | | 1.7666 | 2.6280 | H8 | 2.1512 | 1.0879 | 2.4142 | 2.4839 | 3.0602 | 2.4994 | 1.7666 | | 2.6280 | H9 | 3.7066 | 2.3785 | 1.3310 | 4.5172 | 4.0176 | 4.0176 | 2.6280 | 2.6280 | |
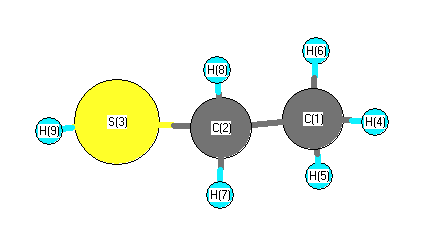 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.130 |
|
C1 |
C2 |
H7 |
110.001 |
| C1 |
C2 |
H8 |
110.001 |
|
C2 |
C1 |
H4 |
110.012 |
| C2 |
C1 |
H5 |
110.985 |
|
C2 |
C1 |
H6 |
110.985 |
| C2 |
S3 |
H9 |
96.729 |
|
S3 |
C2 |
H7 |
109.564 |
| S3 |
C2 |
H8 |
109.564 |
|
H4 |
C1 |
H5 |
108.202 |
| H4 |
C1 |
H6 |
108.202 |
|
H5 |
C1 |
H6 |
108.361 |
| H7 |
C2 |
H8 |
108.569 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -477.198985 |
| Energy at 298.15K | -477.205457 |
| HF Energy | -476.748948 |
| Nuclear repulsion energy | 107.619322 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3234 |
3037 |
22.13 |
|
|
|
| 2 |
A |
3214 |
3019 |
17.94 |
|
|
|
| 3 |
A |
3205 |
3010 |
8.82 |
|
|
|
| 4 |
A |
3151 |
2959 |
17.09 |
|
|
|
| 5 |
A |
3120 |
2930 |
22.91 |
|
|
|
| 6 |
A |
2826 |
2654 |
9.33 |
|
|
|
| 7 |
A |
1544 |
1450 |
2.97 |
|
|
|
| 8 |
A |
1539 |
1445 |
10.07 |
|
|
|
| 9 |
A |
1519 |
1427 |
1.65 |
|
|
|
| 10 |
A |
1462 |
1373 |
4.94 |
|
|
|
| 11 |
A |
1370 |
1286 |
21.88 |
|
|
|
| 12 |
A |
1318 |
1238 |
2.54 |
|
|
|
| 13 |
A |
1168 |
1097 |
12.91 |
|
|
|
| 14 |
A |
1107 |
1040 |
0.63 |
|
|
|
| 15 |
A |
1029 |
966 |
4.99 |
|
|
|
| 16 |
A |
910 |
855 |
8.65 |
|
|
|
| 17 |
A |
771 |
724 |
1.95 |
|
|
|
| 18 |
A |
704 |
661 |
2.41 |
|
|
|
| 19 |
A |
343 |
322 |
2.96 |
|
|
|
| 20 |
A |
284 |
266 |
2.48 |
|
|
|
| 21 |
A |
255 |
239 |
18.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17035.3 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 15999.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.628 |
-0.350 |
-0.055 |
| C2 |
0.491 |
0.647 |
0.090 |
| S3 |
-1.160 |
-0.098 |
-0.074 |
| H4 |
2.589 |
0.161 |
0.005 |
| H5 |
1.596 |
-1.097 |
0.737 |
| H6 |
1.568 |
-0.868 |
-1.010 |
| H7 |
0.559 |
1.176 |
1.039 |
| H8 |
0.538 |
1.394 |
-0.700 |
| H9 |
-1.009 |
-0.983 |
0.908 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5190 | 2.7991 | 1.0899 | 1.0890 | 1.0876 | 2.1601 | 2.1558 | 2.8778 |
C2 | 1.5190 | | 1.8184 | 2.1550 | 2.1635 | 2.1597 | 1.0884 | 1.0888 | 2.3618 | S3 | 2.7991 | 1.8184 | | 3.7583 | 3.0415 | 2.9852 | 2.4116 | 2.3455 | 1.3316 | H4 | 1.0899 | 2.1550 | 3.7583 | | 1.7617 | 1.7689 | 2.4934 | 2.4950 | 3.8818 | H5 | 1.0890 | 2.1635 | 3.0415 | 1.7617 | | 1.7616 | 2.5164 | 3.0647 | 2.6129 | H6 | 1.0876 | 2.1597 | 2.9852 | 1.7689 | 1.7616 | | 3.0644 | 2.5051 | 3.2148 | H7 | 2.1601 | 1.0884 | 2.4116 | 2.4934 | 2.5164 | 3.0644 | | 1.7528 | 2.6720 | H8 | 2.1558 | 1.0888 | 2.3455 | 2.4950 | 3.0647 | 2.5051 | 1.7528 | | 3.2613 | H9 | 2.8778 | 2.3618 | 1.3316 | 3.8818 | 2.6129 | 3.2148 | 2.6720 | 3.2613 | |
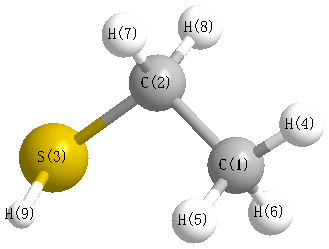 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.711 |
|
C1 |
C2 |
H7 |
110.803 |
| C1 |
C2 |
H8 |
110.430 |
|
C2 |
C1 |
H4 |
110.299 |
| C2 |
C1 |
H5 |
111.035 |
|
C2 |
C1 |
H6 |
110.814 |
| C2 |
S3 |
H9 |
95.906 |
|
S3 |
C2 |
H7 |
109.554 |
| S3 |
C2 |
H8 |
104.764 |
|
H4 |
C1 |
H5 |
107.896 |
| H4 |
C1 |
H6 |
108.645 |
|
H5 |
C1 |
H6 |
108.054 |
| H7 |
C2 |
H8 |
107.229 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
