return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-2688.210721
Energy at 298.15K-2688.221413
HF Energy-2687.607990
Nuclear repulsion energy252.184552
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3239 3042 16.04      
2 A' 3215 3019 29.49      
3 A' 3188 2994 0.86      
4 A' 3118 2928 24.91      
5 A' 1553 1459 8.49      
6 A' 1542 1448 10.05      
7 A' 1468 1379 7.34      
8 A' 1309 1229 48.76      
9 A' 1224 1150 20.14      
10 A' 1095 1028 7.47      
11 A' 932 875 5.77      
12 A' 590 554 14.83      
13 A' 435 408 1.84      
14 A' 322 303 0.73      
15 A' 304 286 1.03      
16 A" 3236 3039 9.36      
17 A" 3209 3014 1.26      
18 A" 3114 2925 13.08      
19 A" 1532 1439 0.44      
20 A" 1529 1436 3.24      
21 A" 1452 1364 11.45      
22 A" 1392 1307 0.53      
23 A" 1185 1113 0.54      
24 A" 985 925 0.08      
25 A" 965 906 1.10      
26 A" 311 292 0.45      
27 A" 275 258 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 21357.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 20059.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.27156 0.09767 0.07717

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.464 -0.894 0.000
Br2 -0.067 1.003 0.000
H3 1.552 -0.866 0.000
C4 -0.067 -1.546 1.259
C5 -0.067 -1.546 -1.259
H6 -1.154 -1.501 1.274
H7 0.238 -2.594 1.276
H8 0.313 -1.059 2.154
H9 -1.154 -1.501 -1.274
H10 0.238 -2.594 -1.276
H11 0.313 -1.059 -2.154

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.96971.08771.51431.51432.14732.13752.16562.14732.13752.1656
Br21.96972.47222.84302.84303.01203.82833.00603.01203.82833.0060
H31.08772.47222.16042.16043.05732.51762.49223.05732.51762.4922
C41.51432.84302.16042.51861.08861.09121.08712.75732.76033.4687
C51.51432.84302.16042.51862.75732.76033.46871.08861.09121.0871
H62.14733.01203.05731.08862.75731.77031.76682.54823.10463.7549
H72.13753.82832.51761.09122.76031.77031.76933.10462.55253.7586
H82.16563.00602.49221.08713.46871.76681.76933.75493.75864.3080
H92.14733.01203.05732.75731.08862.54823.10463.75491.77031.7668
H102.13753.82832.51762.76031.09123.10462.55253.75861.77031.7693
H112.16563.00602.49223.46871.08713.75493.75864.30801.76681.7693

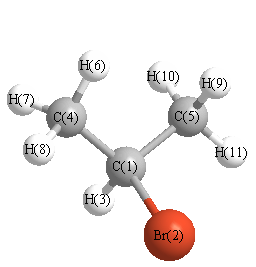
picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.094 C1 C4 H7 109.172
C1 C4 H8 111.657 C1 C5 H9 110.094
C1 C5 H10 109.172 C1 C5 H11 111.657
Br2 C1 H3 104.175 Br2 C1 C4 108.674
Br2 C1 C5 108.674 H3 C1 C4 111.197
H3 C1 C5 111.197 C4 C1 C5 112.527
H6 C4 H7 108.620 H6 C4 H8 108.602
H7 C4 H8 108.634 H9 C5 H10 108.620
H9 C5 H11 108.602 H10 C5 H11 108.634
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability