return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (Propylene oxide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-192.556259
Energy at 298.15K-192.563207
HF Energy-191.922192
Nuclear repulsion energy124.592210
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3297 3097 19.99      
2 A 3234 3037 23.88      
3 A 3216 3020 17.24      
4 A 3207 3012 12.03      
5 A 3193 2999 20.07      
6 A 3121 2931 16.51      
7 A 1577 1481 5.07      
8 A 1546 1452 7.07      
9 A 1528 1435 4.95      
10 A 1487 1396 19.48      
11 A 1443 1355 4.25      
12 A 1319 1239 2.44      
13 A 1220 1145 0.41      
14 A 1194 1121 3.67      
15 A 1181 1109 0.66      
16 A 1148 1078 6.25      
17 A 1065 1000 6.99      
18 A 992 932 11.46      
19 A 930 873 1.90      
20 A 869 816 48.52      
21 A 784 736 6.59      
22 A 415 390 4.37      
23 A 373 350 5.06      
24 A 218 205 0.57      

Unscaled Zero Point Vibrational Energy (zpe) 19276.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 18104.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.59894 0.22359 0.19852

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.829 -0.798 -0.234
C2 -1.502 0.094 -0.151
H3 -1.391 0.307 -1.212
H4 -2.060 -0.834 -0.041
H5 -2.075 0.898 0.312
C6 -0.155 -0.027 0.493
H7 -0.151 -0.235 1.558
C8 1.033 0.624 -0.066
H9 0.934 1.205 -0.975
H10 1.860 0.891 0.578

Atom - Atom Distances (Å)
  O1 C2 H3 H4 H5 C6 H7 C8 H9 H10
O12.49732.66492.89563.40671.44572.11891.44652.13812.1397
C22.49731.08781.08871.08991.49892.20422.59122.80143.5320
H32.66491.08781.76671.77232.13253.08342.69942.50353.7572
H42.89561.08871.76671.76722.13752.56193.41953.74114.3281
H53.40671.08991.77231.76722.13942.55743.14273.28733.9447
C61.44571.49892.13252.13752.13941.08561.46482.20392.2159
H72.11892.20423.08342.56192.55741.08562.18593.10952.5051
C81.44652.59122.69943.41953.14271.46482.18591.08311.0824
H92.13812.80142.50353.74113.28732.20393.10951.08311.8353
H102.13973.53203.75724.32813.94472.21592.50511.08241.8353

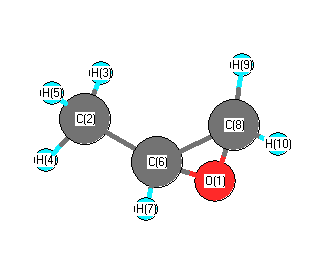
picture of Propylene oxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C6 C2 116.006 O1 C6 H7 112.901
O1 C6 C8 59.598 O1 C8 C6 59.543
O1 C8 H9 114.634 O1 C8 H10 114.823
C2 C6 H7 116.128 C2 C6 C8 121.922
H3 C2 H4 108.525 H3 C2 H5 108.949
H3 C2 C6 110.039 H4 C2 H5 108.420
H4 C2 C6 110.388 H5 C2 C6 110.468
C6 O1 C8 60.859 C6 C8 H9 119.003
C6 C8 H10 120.157 H7 C6 C8 117.203
H9 C8 H10 115.886
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability