Jump to
S1C2
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -637.608891 |
| Energy at 298.15K | -637.613833 |
| HF Energy | -636.991327 |
| Nuclear repulsion energy | 157.171757 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3188 |
2994 |
15.99 |
|
|
|
| 2 |
A' |
3169 |
2976 |
14.16 |
|
|
|
| 3 |
A' |
1569 |
1473 |
0.82 |
|
|
|
| 4 |
A' |
1541 |
1447 |
5.53 |
|
|
|
| 5 |
A' |
1457 |
1368 |
3.88 |
|
|
|
| 6 |
A' |
1339 |
1257 |
16.88 |
|
|
|
| 7 |
A' |
1122 |
1054 |
14.48 |
|
|
|
| 8 |
A' |
1053 |
989 |
111.40 |
|
|
|
| 9 |
A' |
823 |
773 |
39.77 |
|
|
|
| 10 |
A' |
392 |
369 |
2.97 |
|
|
|
| 11 |
A' |
248 |
233 |
13.42 |
|
|
|
| 12 |
A" |
3269 |
3071 |
8.35 |
|
|
|
| 13 |
A" |
3242 |
3045 |
8.58 |
|
|
|
| 14 |
A" |
1333 |
1252 |
0.03 |
|
|
|
| 15 |
A" |
1240 |
1165 |
0.91 |
|
|
|
| 16 |
A" |
1108 |
1041 |
2.58 |
|
|
|
| 17 |
A" |
819 |
769 |
0.32 |
|
|
|
| 18 |
A" |
136 |
128 |
11.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13523.7 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12701.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.611 |
0.000 |
| C2 |
0.970 |
-0.546 |
0.000 |
| Cl3 |
-1.663 |
-0.023 |
0.000 |
| F4 |
2.272 |
-0.014 |
0.000 |
| H5 |
0.136 |
1.222 |
0.887 |
| H6 |
0.136 |
1.222 |
-0.887 |
| H7 |
0.862 |
-1.159 |
0.891 |
| H8 |
0.862 |
-1.159 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5102 | 1.7796 | 2.3566 | 1.0853 | 1.0853 | 2.1616 | 2.1616 |
C2 | 1.5102 | | 2.6840 | 1.4068 | 2.1464 | 2.1464 | 1.0871 | 1.0871 | Cl3 | 1.7796 | 2.6840 | | 3.9348 | 2.3605 | 2.3605 | 2.9086 | 2.9086 | F4 | 2.3566 | 1.4068 | 3.9348 | | 2.6222 | 2.6222 | 2.0233 | 2.0233 | H5 | 1.0853 | 2.1464 | 2.3605 | 2.6222 | | 1.7744 | 2.4888 | 3.0590 | H6 | 1.0853 | 2.1464 | 2.3605 | 2.6222 | 1.7744 | | 3.0590 | 2.4888 | H7 | 2.1616 | 1.0871 | 2.9086 | 2.0233 | 2.4888 | 3.0590 | | 1.7828 | H8 | 2.1616 | 1.0871 | 2.9086 | 2.0233 | 3.0590 | 2.4888 | 1.7828 | |
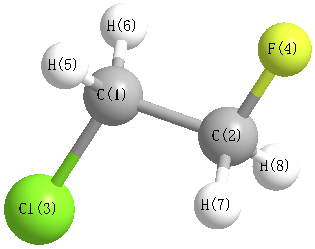 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.728 |
|
C1 |
C2 |
H7 |
111.625 |
| C1 |
C2 |
H8 |
111.625 |
|
C2 |
C1 |
Cl3 |
109.072 |
| C2 |
C1 |
H5 |
110.509 |
|
C2 |
C1 |
H6 |
110.509 |
| Cl3 |
C1 |
H5 |
108.516 |
|
Cl3 |
C1 |
H6 |
108.516 |
| F4 |
C2 |
H7 |
107.758 |
|
F4 |
C2 |
H8 |
107.758 |
| H5 |
C1 |
H6 |
109.666 |
|
H7 |
C2 |
H8 |
110.161 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -637.607930 |
| Energy at 298.15K | -637.612987 |
| HF Energy | -636.989713 |
| Nuclear repulsion energy | 161.422993 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3247 |
3049 |
5.80 |
|
|
|
| 2 |
A |
3225 |
3029 |
13.28 |
|
|
|
| 3 |
A |
3168 |
2975 |
17.43 |
|
|
|
| 4 |
A |
3148 |
2956 |
21.83 |
|
|
|
| 5 |
A |
1551 |
1457 |
1.39 |
|
|
|
| 6 |
A |
1512 |
1420 |
9.89 |
|
|
|
| 7 |
A |
1457 |
1368 |
15.73 |
|
|
|
| 8 |
A |
1394 |
1309 |
33.14 |
|
|
|
| 9 |
A |
1302 |
1223 |
0.67 |
|
|
|
| 10 |
A |
1249 |
1173 |
0.93 |
|
|
|
| 11 |
A |
1135 |
1066 |
28.05 |
|
|
|
| 12 |
A |
1088 |
1021 |
55.11 |
|
|
|
| 13 |
A |
1012 |
951 |
10.26 |
|
|
|
| 14 |
A |
876 |
823 |
14.72 |
|
|
|
| 15 |
A |
731 |
687 |
24.65 |
|
|
|
| 16 |
A |
476 |
447 |
13.34 |
|
|
|
| 17 |
A |
293 |
276 |
0.93 |
|
|
|
| 18 |
A |
140 |
131 |
2.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13500.5 cm
-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12679.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31+G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.096 |
0.843 |
-0.298 |
| C2 |
1.191 |
0.429 |
0.362 |
| Cl3 |
-1.419 |
-0.287 |
0.066 |
| F4 |
1.654 |
-0.767 |
-0.201 |
| H5 |
-0.384 |
1.828 |
0.063 |
| H6 |
0.032 |
0.877 |
-1.376 |
| H7 |
1.954 |
1.187 |
0.188 |
| H8 |
1.063 |
0.263 |
1.430 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5044 | 1.7770 | 2.3805 | 1.0880 | 1.0861 | 2.1352 | 2.1599 |
C2 | 1.5044 | | 2.7219 | 1.4014 | 2.1275 | 2.1364 | 1.0899 | 1.0878 | Cl3 | 1.7770 | 2.7219 | | 3.1215 | 2.3544 | 2.3534 | 3.6829 | 2.8846 | F4 | 2.3805 | 1.4014 | 3.1215 | | 3.3108 | 2.5916 | 2.0154 | 2.0171 | H5 | 1.0880 | 2.1275 | 2.3544 | 3.3108 | | 1.7745 | 2.4278 | 2.5321 | H6 | 1.0861 | 2.1364 | 2.3534 | 2.5916 | 1.7745 | | 2.4975 | 3.0515 | H7 | 2.1352 | 1.0899 | 3.6829 | 2.0154 | 2.4278 | 2.4975 | | 1.7859 | H8 | 2.1599 | 1.0878 | 2.8846 | 2.0171 | 2.5321 | 3.0515 | 1.7859 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.969 |
|
C1 |
C2 |
H7 |
109.752 |
| C1 |
C2 |
H8 |
111.869 |
|
C2 |
C1 |
Cl3 |
111.828 |
| C2 |
C1 |
H5 |
109.251 |
|
C2 |
C1 |
H6 |
110.066 |
| Cl3 |
C1 |
H5 |
108.106 |
|
Cl3 |
C1 |
H6 |
108.128 |
| F4 |
C2 |
H7 |
107.339 |
|
F4 |
C2 |
H8 |
107.600 |
| H5 |
C1 |
H6 |
109.411 |
|
H7 |
C2 |
H8 |
110.188 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
