return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-235.094320
Energy at 298.15K-235.106699
HF Energy-234.198901
Nuclear repulsion energy231.911741
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3227 3031 60.62      
2 A 3217 3021 2.86      
3 A 3211 3016 51.36      
4 A 3189 2995 1.66      
5 A 3117 2927 7.49      
6 A 3115 2925 1.28      
7 A 1731 1626 2.03      
8 A 1551 1457 10.20      
9 A 1544 1450 3.51      
10 A 1538 1445 0.15      
11 A 1451 1362 0.17      
12 A 1379 1295 0.14      
13 A 1329 1248 0.08      
14 A 1300 1221 0.37      
15 A 1117 1050 0.61      
16 A 1098 1031 0.49      
17 A 1046 982 0.62      
18 A 991 930 0.57      
19 A 870 817 0.54      
20 A 794 746 0.51      
21 A 523 491 0.14      
22 A 324 304 0.00      
23 A 217 204 0.01      
24 A 197 185 0.00      
25 A 42 39 0.01      
26 B 3216 3021 66.02      
27 B 3211 3015 3.92      
28 B 3202 3007 6.78      
29 B 3176 2983 5.10      
30 B 3114 2925 73.40      
31 B 3114 2924 34.82      
32 B 1548 1454 4.89      
33 B 1544 1450 8.25      
34 B 1524 1431 1.14      
35 B 1469 1380 4.00      
36 B 1449 1361 5.21      
37 B 1361 1278 8.63      
38 B 1315 1235 0.01      
39 B 1205 1132 0.22      
40 B 1124 1055 7.20      
41 B 1075 1010 2.15      
42 B 936 879 5.42      
43 B 828 777 4.03      
44 B 766 720 44.04      
45 B 565 530 4.30      
46 B 364 342 1.26      
47 B 246 231 0.08      
48 B 37 35 0.12      

Unscaled Zero Point Vibrational Energy (zpe) 37251.7 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 34986.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.27301 0.05210 0.04959

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.007 0.673 0.877
C2 -0.007 -0.673 0.877
C3 -0.007 1.583 -0.314
C4 0.007 -1.583 -0.314
C5 -1.316 2.369 -0.410
C6 1.316 -2.369 -0.410
H7 0.001 1.172 1.842
H8 -0.001 -1.172 1.842
H9 0.153 1.021 -1.233
H10 -0.153 -1.021 -1.233
H11 0.825 2.287 -0.231
H12 -0.825 -2.287 -0.231
H13 -1.301 3.059 -1.253
H14 -2.159 1.691 -0.537
H15 -1.486 2.949 0.497
H16 1.301 -3.059 -1.253
H17 2.159 -1.691 -0.537
H18 1.486 -2.949 0.497

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34601.49892.55122.50683.55331.08642.08222.14312.71032.12183.26813.45572.77902.74814.48783.49503.9304
C21.34602.55121.49893.55332.50682.08221.08642.71032.14313.26812.12184.48783.49503.93043.45572.77902.7481
C31.49892.55123.16671.52994.16932.19453.49851.08912.76561.09303.95672.17592.16582.17024.91383.93194.8399
C42.55121.49893.16674.16931.52993.49852.19452.76561.08913.95671.09304.91383.93194.83992.17592.16582.1702
C52.50683.55331.52994.16935.42052.87054.39802.15673.67762.15054.68541.08951.08911.08976.08495.34556.0791
C63.55332.50684.16931.52995.42054.39802.87053.67762.15674.68542.15056.08495.34556.07911.08951.08911.0897
H71.08642.08222.19453.49852.87054.39802.34433.08203.77982.49364.11633.85173.25422.67895.40124.30214.5819
H82.08221.08643.49852.19454.39802.87052.34433.77983.08204.11632.49365.40124.30214.58193.85173.25422.6789
H92.14312.71031.08912.76562.15673.67763.08203.77982.06471.74903.59202.50332.50503.06484.23873.44444.5308
H102.71032.14312.76561.08913.67762.15673.77983.08202.06473.59201.74904.23873.44444.53082.50332.50503.0648
H112.12183.26811.09303.95672.15054.68542.49364.11631.74903.59204.86252.48243.05822.51235.46364.20645.3269
H123.26812.12183.95671.09304.68542.15054.11632.49363.59201.74904.86255.46364.20645.32692.48243.05822.5123
H133.45574.48782.17594.91381.08956.08493.85175.40122.50334.23872.48245.46361.76691.76366.64845.91986.8500
H142.77903.49502.16583.93191.08915.34553.25424.30212.50503.44443.05824.20641.76691.76115.91985.48405.9898
H152.74813.93042.17024.83991.08976.07912.67894.58193.06484.53082.51235.32691.76361.76116.85005.98986.6039
H164.48783.45574.91382.17596.08491.08955.40123.85174.23872.50335.46362.48246.64845.91986.85001.76691.7636
H173.49502.77903.93192.16585.34551.08914.30213.25423.44442.50504.20643.05825.91985.48405.98981.76691.7611
H183.93042.74814.83992.17026.07911.08974.58192.67894.53083.06485.32692.51236.85005.98986.60391.76361.7611

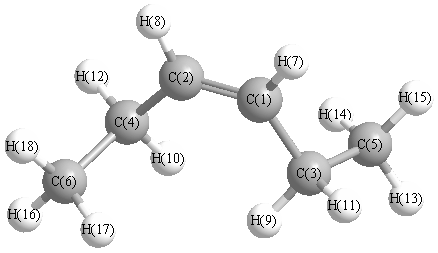
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.391 C1 C2 H8 117.349
C1 C3 C5 111.714 C1 C3 H9 110.813
C1 C3 H11 108.885 C2 C1 C3 127.391
C2 C1 H7 117.349 C2 C4 C6 111.714
C2 C4 H10 110.813 C2 C4 H12 108.885
C3 C1 H7 115.239 C3 C5 H13 111.222
C3 C5 H14 110.442 C3 C5 H15 110.755
C4 C2 H8 115.239 C4 C6 H16 111.222
C4 C6 H17 110.442 C4 C6 H18 110.755
C5 C3 H9 109.719 C5 C3 H11 109.008
C6 C4 H10 109.719 C6 C4 H12 109.008
H9 C3 H11 106.548 H10 C4 H12 106.548
H13 C5 H14 108.392 H13 C5 H15 108.057
H14 C5 H15 107.859 H16 C6 H17 108.392
H16 C6 H18 108.057 H17 C6 H18 107.859
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability