return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: CCSD/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCSD/6-31+G**
 hartrees
Energy at 0K-871.757640
Energy at 298.15K-871.765422
HF Energy-871.400727
Nuclear repulsion energy192.701472
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2295 2143 168.02      
2 A1 2285 2134 29.89      
3 A1 2261 2111 99.57      
4 A1 987 921 83.18      
5 A1 964 900 20.70      
6 A1 932 870 215.20      
7 A1 588 549 10.38      
8 A1 402 376 1.00      
9 A1 102 95 2.10      
10 A2 2290 2139 0.00      
11 A2 976 912 0.00      
12 A2 729 681 0.00      
13 A2 431 402 0.00      
14 A2 80 75 0.00      
15 B1 2295 2143 302.82      
16 B1 2271 2120 44.49      
17 B1 980 915 97.87      
18 B1 621 580 11.91      
19 B1 325 303 31.48      
20 B1 100 93 0.07      
21 B2 2294 2142 104.43      
22 B2 2279 2129 112.90      
23 B2 977 912 37.86      
24 B2 923 862 367.43      
25 B2 759 709 350.42      
26 B2 486 454 5.74      
27 B2 455 425 28.54      

Unscaled Zero Point Vibrational Energy (zpe) 15042.8 cm-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 14047.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD/6-31+G**
ABC
0.31463 0.06565 0.05755

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.885
Si2 0.000 1.948 -0.417
Si3 0.000 -1.948 -0.417
H4 1.198 0.000 1.759
H5 -1.198 0.000 1.759
H6 0.000 3.153 0.439
H7 0.000 -3.153 0.439
H8 1.200 1.978 -1.280
H9 -1.200 1.978 -1.280
H10 -1.200 -1.978 -1.280
H11 1.200 -1.978 -1.280

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.34292.34291.48221.48223.18493.18493.16943.16943.16943.1694
Si22.34293.89523.15613.15611.47845.17221.47911.47914.19514.1951
Si32.34293.89523.15613.15615.17221.47844.19514.19511.47911.4791
H41.48223.15613.15612.39513.62233.62233.62644.34754.34753.6264
H51.48223.15613.15612.39513.62233.62234.34753.62643.62644.3475
H63.18491.47845.17223.62233.62236.30672.40362.40365.54345.5434
H73.18495.17221.47843.62233.62236.30675.54345.54342.40362.4036
H83.16941.47914.19513.62644.34752.40365.54342.40064.62783.9565
H93.16941.47914.19514.34753.62642.40365.54342.40063.95654.6278
H103.16944.19511.47914.34753.62645.54342.40364.62783.95652.4006
H113.16944.19511.47913.62644.34755.54342.40363.95654.62782.4006

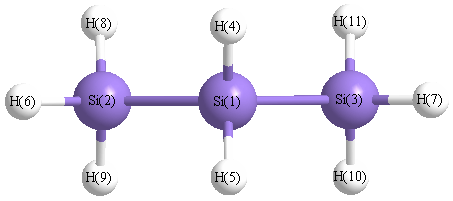
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 110.878 S1 S2 H8 109.985
S1 S2 H9 109.985 S1 S3 H7 110.878
S1 S3 H10 109.985 S1 S3 H11 109.985
S2 S1 S3 112.465 S2 S1 H4 109.119
S2 S1 H5 109.119 S3 S1 H4 109.119
S3 S1 H5 109.119 H4 S1 H5 107.788
H6 S2 H8 108.724 H6 S2 H9 108.724
H7 S3 H10 108.724 H7 S3 H11 108.724
H8 S2 H9 108.493 H10 S3 H11 108.493
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability