return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Butane, 2-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-197.124182
Energy at 298.15K-197.137202
HF Energy-196.351361
Nuclear repulsion energy193.153939
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3222 3026 30.33      
2 A 3212 3017 41.97      
3 A 3205 3010 40.36      
4 A 3204 3009 29.05      
5 A 3197 3003 57.28      
6 A 3193 2999 3.74      
7 A 3149 2958 24.68      
8 A 3116 2926 25.10      
9 A 3106 2918 30.20      
10 A 3103 2914 39.31      
11 A 3096 2907 36.62      
12 A 3087 2899 7.88      
13 A 1561 1466 3.07      
14 A 1552 1457 16.05      
15 A 1548 1454 5.40      
16 A 1546 1452 9.68      
17 A 1540 1446 2.93      
18 A 1533 1440 0.00      
19 A 1524 1432 0.10      
20 A 1461 1372 6.81      
21 A 1456 1367 4.03      
22 A 1443 1356 7.61      
23 A 1422 1335 0.68      
24 A 1401 1316 0.87      
25 A 1355 1273 1.79      
26 A 1327 1246 1.62      
27 A 1240 1165 0.14      
28 A 1231 1156 0.61      
29 A 1206 1133 1.70      
30 A 1092 1026 0.18      
31 A 1059 995 1.76      
32 A 1007 946 2.15      
33 A 995 934 0.26      
34 A 954 896 0.54      
35 A 950 893 0.70      
36 A 822 772 0.19      
37 A 789 741 1.50      
38 A 466 438 0.11      
39 A 426 400 0.07      
40 A 376 353 0.04      
41 A 284 267 0.01      
42 A 266 250 0.01      
43 A 234 220 0.00      
44 A 221 208 0.03      
45 A 98 92 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 36136.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 33939.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.24635 0.11302 0.08635

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.916 -0.616 0.282
H2 -2.184 0.020 -1.189
H3 -2.168 0.973 0.294
C4 -2.079 -0.036 -0.105
H5 -0.759 -1.726 -0.038
H6 -0.640 -0.683 1.364
C7 -0.750 -0.679 0.275
H8 0.352 -0.007 -1.424
C9 0.479 -0.004 -0.336
H10 -0.217 2.060 -0.184
H11 1.530 1.890 -0.296
H12 0.704 1.491 1.210
C13 0.627 1.445 0.121
H14 2.621 -0.349 -0.467
H15 1.662 -1.827 -0.348
H16 1.904 -0.813 1.076
C17 1.741 -0.795 -0.003

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 H6 C7 H8 C9 H10 H11 H12 C13 H14 H15 H16 C17
H11.76131.75681.08932.44682.52092.16713.73673.50493.82885.13674.28994.10225.59334.77664.88894.6689
H21.76131.76251.09052.53093.06522.16542.54762.79713.00664.25364.03213.41354.87274.34844.74724.1805
H31.75681.76251.08833.06292.49432.17723.20372.89122.28323.85563.05852.84005.02574.78714.51464.3005
C41.08931.09051.08832.14602.15571.52482.76662.56922.80424.09573.43563.09324.72424.15464.22653.8961
H52.44682.53093.06292.14601.75161.09302.47212.14193.82694.28773.74733.46403.67422.44233.02742.6674
H62.52093.06522.49432.15571.75161.09473.03602.14583.17733.75352.56002.77083.75503.08862.56402.7483
C72.16712.16542.17721.52481.09301.09472.13391.52952.82713.48212.77362.53553.46692.74242.77532.5088
H83.73672.54763.20372.76662.47213.03602.13391.09602.47692.50213.05042.13832.48552.48663.05072.1373
C93.50492.79712.89122.56922.14192.14581.52951.09602.18312.16642.16141.52592.17292.17272.16231.5258
H103.82883.00662.28322.80423.82693.17732.82712.47692.18311.75941.76471.08853.73324.31973.78623.4666
H115.13674.25363.85564.09574.28773.75353.48212.50212.16641.75941.76371.09012.49673.71953.05402.7097
H124.28994.03213.05853.43563.74732.56002.77363.05042.16141.76471.76371.09213.14193.78772.60052.7879
C134.10223.41352.84003.09323.46402.77082.53552.13831.52591.08851.09011.09212.74543.46272.76312.5045
H145.59334.87275.02574.72423.67423.75503.46692.48552.17293.73322.49673.14192.74541.76521.76331.0901
H154.77664.34844.78714.15462.44233.08862.74242.48662.17274.31973.71953.78773.46271.76521.76461.0901
H164.88894.74724.51464.22653.02742.56402.77533.05072.16233.78623.05402.60052.76311.76331.76461.0916
C174.66894.18054.30053.89612.66742.74832.50882.13731.52583.46662.70972.78792.50451.09011.09011.0916

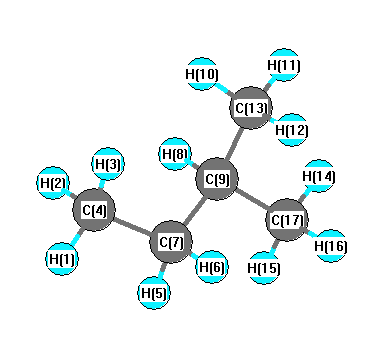
picture of Butane, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 107.806 H1 C4 H3 107.558
H1 C4 C7 110.889 H2 C4 H3 107.983
H2 C4 C7 110.678 H3 C4 C7 111.765
C4 C7 H5 109.004 C4 C7 H6 109.666
C4 C7 C9 114.527 H5 C7 H6 106.384
H5 C7 C9 108.369 H6 C7 C9 108.574
C7 C9 H8 107.579 C7 C9 C13 112.160
C7 C9 C17 110.396 H8 C9 C13 108.160
H8 C9 C17 108.092 C9 C13 H10 112.156
C9 C13 H11 110.712 C9 C13 H12 110.199
C9 C17 H14 111.237 C9 C17 H15 111.222
C9 C17 H16 110.308 H10 C13 H11 107.720
H10 C13 H12 108.051 H11 C13 H12 107.851
C13 C9 C17 110.306 H14 C17 H15 108.129
H14 C17 H16 107.848 H15 C17 H16 107.965
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability