return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Butadiene, 2-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-194.695857
Energy at 298.15K-194.703376
HF Energy-193.976331
Nuclear repulsion energy160.444685
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3341 3138 11.17      
2 A' 3327 3125 12.08      
3 A' 3245 3048 3.91      
4 A' 3229 3032 13.25      
5 A' 3225 3029 10.92      
6 A' 3225 3029 11.91      
7 A' 3114 2925 19.39      
8 A' 1725 1620 2.49      
9 A' 1675 1573 16.86      
10 A' 1550 1456 7.74      
11 A' 1502 1410 0.82      
12 A' 1471 1381 2.26      
13 A' 1448 1360 5.91      
14 A' 1366 1283 0.61      
15 A' 1346 1264 0.36      
16 A' 1110 1043 3.01      
17 A' 1031 969 0.20      
18 A' 990 930 0.56      
19 A' 812 762 0.18      
20 A' 537 504 0.10      
21 A' 425 399 1.28      
22 A' 284 267 0.68      
23 A" 3198 3003 13.56      
24 A" 1518 1426 8.49      
25 A" 1081 1015 1.93      
26 A" 1013 951 25.24      
27 A" 893 839 47.68      
28 A" 880 826 45.22      
29 A" 762 716 1.11      
30 A" 627 589 1.01      
31 A" 360 338 9.66      
32 A" 196 184 0.62      
33 A" 122 115 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 25312.5 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23773.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.28566 0.13927 0.09527

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.589 1.736 0.000
C2 0.000 0.523 0.000
C3 -0.829 -0.684 0.000
C4 -0.363 -1.947 0.000
C5 1.495 0.373 0.000
H6 -1.666 1.836 0.000
H7 -0.008 2.648 0.000
H8 -1.902 -0.525 0.000
H9 0.694 -2.170 0.000
H10 -1.041 -2.787 0.000
H11 1.979 1.347 0.000
H12 1.830 -0.178 0.879
H13 1.830 -0.178 -0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34872.43273.69022.49041.08171.08102.61514.11194.54622.59763.20743.2074
C21.34871.46472.49651.50252.12152.12462.17212.78133.47032.14382.14762.1476
C32.43271.46471.34582.55332.65613.43191.08532.12782.11353.46592.84582.8458
C43.69022.49651.34582.97224.00144.60842.09581.08041.08004.04162.95132.9513
C52.49041.50252.55332.97223.48342.72623.51422.66634.05231.08761.09021.0902
H61.08172.12152.65614.00143.48341.84632.37314.65014.66563.67804.12934.1293
H71.08102.12463.43194.60842.72621.84633.69544.86875.53242.37483.48323.4832
H82.61512.17211.08532.09583.51422.37313.69543.07402.42064.30943.85013.8501
H94.11192.78132.12781.08042.66634.65014.86873.07401.84203.74452.45602.4560
H104.54623.47032.11351.08004.05234.66565.53242.42061.84205.12013.97813.9781
H112.59762.14383.46594.04161.08763.67802.37484.30943.74455.12011.76641.7664
H123.20742.14762.84582.95131.09024.12933.48323.85012.45603.97811.76641.7585
H133.20742.14762.84582.95131.09024.12933.48323.85012.45603.97811.76641.7585

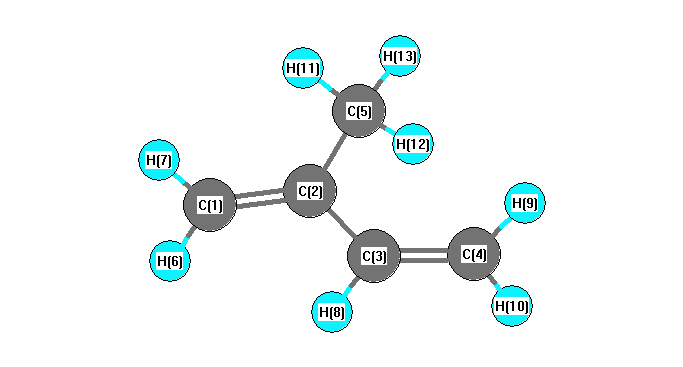
picture of 1,3-Butadiene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.633 C1 C2 C5 121.627
C2 C1 H6 121.199 C2 C1 H7 121.570
C2 C3 C4 125.267 C2 C3 H8 116.020
C2 C5 H11 110.703 C2 C5 H12 110.851
C2 C5 H13 110.851 C3 C2 C5 118.740
C3 C4 H9 122.188 C3 C4 H10 120.817
C4 C3 H8 118.713 H6 C1 H7 117.231
H9 C4 H10 116.994 H11 C5 H12 108.409
H11 C5 H13 108.409 H12 C5 H13 107.509
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability