return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH(CH3)CH3 (Propanal, 2-methyl-)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-231.781492
Energy at 298.15K-231.790016
HF Energy-231.001482
Nuclear repulsion energy175.600491
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3219 3023 23.20      
2 A' 3210 3015 43.38      
3 A' 3156 2964 5.18      
4 A' 3113 2924 18.19      
5 A' 2977 2796 93.15      
6 A' 1762 1654 146.59      
7 A' 1555 1461 18.17      
8 A' 1549 1455 7.44      
9 A' 1463 1374 3.45      
10 A' 1438 1351 1.20      
11 A' 1351 1269 0.99      
12 A' 1218 1144 0.25      
13 A' 1211 1138 10.41      
14 A' 963 905 15.92      
15 A' 871 818 15.44      
16 A' 551 518 5.09      
17 A' 358 336 8.08      
18 A' 338 318 0.13      
19 A' 242 227 1.02      
20 A" 3219 3023 12.35      
21 A" 3206 3011 3.39      
22 A" 3112 2923 24.65      
23 A" 1537 1443 2.38      
24 A" 1532 1439 1.25      
25 A" 1440 1352 4.23      
26 A" 1377 1294 0.67      
27 A" 1169 1098 1.34      
28 A" 1017 955 0.76      
29 A" 975 915 0.03      
30 A" 953 895 1.23      
31 A" 337 317 0.89      
32 A" 216 203 0.02      
33 A" 63 59 7.18      

Unscaled Zero Point Vibrational Energy (zpe) 25347.6 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 23806.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.25881 0.12399 0.09396

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.229 0.595 2.162
H2 -0.229 0.595 -2.162
H3 -1.696 0.112 -1.303
H4 -1.696 0.112 1.303
H5 -1.165 1.792 1.264
H6 -1.165 1.792 -1.264
C7 -0.819 0.759 -1.262
C8 -0.819 0.759 1.262
H9 0.904 1.094 0.000
C10 0.000 0.483 0.000
C11 0.440 -0.954 0.000
H12 -0.384 -1.694 0.000
O13 1.605 -1.336 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 C10 C11 H12 O13
H14.32493.79401.76731.76533.74873.47881.08902.49152.17742.74293.15293.4303
H24.32491.76733.79403.74871.76531.08903.47882.49152.17742.74293.15293.4303
H33.79401.76732.60613.11381.76231.09092.78713.06902.17052.71962.58443.8327
H41.76733.79402.60611.76233.11382.78711.09093.06902.17052.71962.58443.8327
H51.76533.74873.11381.76232.52862.75111.08912.52282.16073.42313.78974.3654
H63.74871.76531.76233.11382.52861.08912.75112.52282.16073.42313.78974.3654
C73.47881.08901.09092.78712.75111.08912.52402.16131.52952.47282.79333.4436
C81.08903.47882.78711.09091.08912.75112.52402.16131.52952.47282.79333.4436
H92.49152.49153.06903.06902.52282.52282.16132.16131.09082.10043.07172.5294
C102.17742.17742.17052.17052.16072.16071.52951.52951.09081.50352.21132.4261
C112.74292.74292.71962.71963.42313.42312.47282.47282.10041.50351.10801.2260
H123.15293.15292.58442.58443.78973.78972.79332.79333.07172.21131.10802.0218
O133.43033.43033.83273.83274.36544.36543.44363.44362.52942.42611.22602.0218

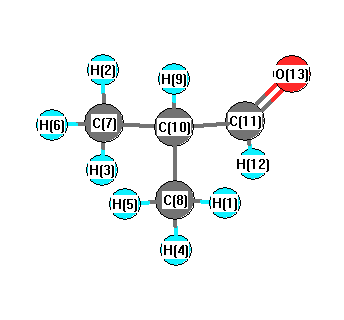
picture of Propanal, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C8 H4 108.337 H1 C8 H5 108.292
H1 C8 C10 111.405 H2 C7 H3 108.337
H2 C7 H6 108.292 H2 C7 C10 111.405
H3 C7 H6 107.885 H3 C7 C10 110.740
H4 C8 H5 107.885 H4 C8 C10 110.740
H5 C8 C10 110.073 H6 C7 C10 110.073
C7 C10 C8 111.204 C7 C10 H9 110.015
C7 C10 C11 109.232 C8 C10 H9 110.015
C8 C10 C11 109.232 H9 C10 C11 107.044
C10 C11 H12 114.878 C10 C11 O13 125.152
H12 C11 O13 119.970
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability