return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H10N2 (1,2-Diaminopropane)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-229.157415
Energy at 298.15K-229.170045
HF Energy-228.335391
Nuclear repulsion energy195.557153
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3701 3476 4.14      
2 A 3673 3450 2.13      
3 A 3582 3364 0.43      
4 A 3570 3353 0.16      
5 A 3220 3024 22.99      
6 A 3207 3012 26.54      
7 A 3186 2993 21.71      
8 A 3119 2929 42.31      
9 A 3113 2923 14.55      
10 A 3023 2839 70.02      
11 A 1698 1595 3.89      
12 A 1688 1585 78.06      
13 A 1547 1453 4.45      
14 A 1539 1446 6.64      
15 A 1521 1428 0.95      
16 A 1457 1368 4.85      
17 A 1446 1358 9.73      
18 A 1425 1338 11.48      
19 A 1402 1316 8.65      
20 A 1373 1290 7.37      
21 A 1288 1209 0.81      
22 A 1237 1162 1.78      
23 A 1200 1127 4.29      
24 A 1142 1072 18.93      
25 A 1070 1005 8.19      
26 A 1052 988 1.57      
27 A 986 926 3.44      
28 A 918 862 54.05      
29 A 895 840 77.83      
30 A 856 803 100.27      
31 A 799 750 133.17      
32 A 505 475 1.52      
33 A 483 454 19.80      
34 A 380 357 1.29      
35 A 351 329 56.46      
36 A 269 252 27.58      
37 A 257 242 13.72      
38 A 233 219 22.09      
39 A 134 125 4.80      

Unscaled Zero Point Vibrational Energy (zpe) 31270.2 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 29369.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.26769 0.11707 0.08947

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.496 1.381 -0.212
H2 -0.256 1.946 0.168
H3 1.362 1.847 0.035
N4 -2.050 -0.126 0.028
H5 -2.158 0.751 -0.465
H6 -2.212 0.031 1.016
C7 -0.741 -0.719 -0.219
H8 -0.748 -1.732 0.190
H9 -0.616 -0.799 -1.300
C10 1.756 -0.686 -0.029
H11 1.750 -1.713 0.334
H12 1.870 -0.696 -1.113
H13 2.622 -0.182 0.401
C14 0.467 0.026 0.345
H15 0.374 0.031 1.442

Atom - Atom Distances (Å)
  N1 H2 H3 N4 H5 H6 C7 H8 H9 C10 H11 H12 H13 C14 H15
N11.01401.01452.96852.73953.26572.43773.37612.67842.42773.38282.64842.70921.46572.1390
H21.01401.62652.74432.33372.86552.73633.71003.13373.31814.17533.62463.58632.05872.3845
H31.01451.62653.94203.72094.12763.32824.15763.56382.56443.59352.83632.41652.05322.5011
N42.96852.74433.94201.01201.01331.45852.07382.06793.84744.12934.12234.68692.54132.8108
H52.73952.33373.72091.01201.64742.05692.92962.34124.19214.68844.32894.94642.84113.2507
H63.26572.86554.12761.01331.64742.06252.43622.93334.16554.38224.66094.87722.76172.6211
C72.43772.73633.32821.45852.05692.06251.09201.09102.50432.73822.75953.46131.52732.1370
H83.37613.71004.15762.07382.92962.43621.09201.76282.72202.50173.10173.71432.14192.4356
H92.67843.13373.56382.06792.34122.93331.09101.76282.69303.01682.49473.70892.13543.0313
C102.42773.31812.56443.84744.19214.16552.50432.72202.69301.08921.08951.09021.51932.1418
H113.38284.17533.59354.12934.68844.38222.73822.50173.01681.08921.77231.76242.16082.4819
H122.64843.62462.83634.12234.32894.66092.75953.10172.49471.08951.77231.76702.14833.0484
H132.70923.58632.41654.68694.94644.87723.46133.71433.70891.09021.76241.76702.16552.4857
C141.46572.05872.05322.54132.84112.76171.52732.14192.13541.51932.16082.14832.16551.1009
H152.13902.38452.50112.81083.25072.62112.13702.43563.03132.14182.48193.04842.48571.1009

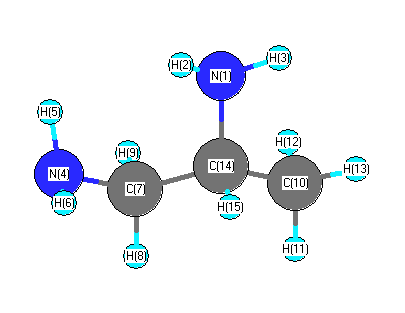
picture of 1,2-Diaminopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 C7 109.048 N1 C14 C10 108.820
N1 C14 H15 112.117 H2 N1 H3 106.612
H2 N1 C14 110.928 H3 N1 C14 110.433
N4 C7 H8 107.938 N4 C7 H9 107.529
N4 C7 C14 116.652 H5 N4 H6 108.861
H5 N4 C7 111.446 H6 N4 C7 111.848
C7 C14 C10 110.565 C7 C14 H15 107.692
H8 C7 H9 107.706 H8 C7 C14 108.576
H9 C7 C14 108.123 C10 C14 H15 108.601
H11 C10 H12 108.863 H11 C10 H13 107.929
H11 C10 C14 110.782 H12 C10 H13 108.318
H12 C10 C14 109.771 H13 C10 C14 111.102
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability