return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCONH2 (Acrylamide)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-246.616219
Energy at 298.15K-246.621954
HF Energy-245.840380
Nuclear repulsion energy162.407727
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3806 3574 49.83      
2 A 3659 3436 58.48      
3 A 3354 3150 1.59      
4 A 3255 3057 10.43      
5 A 3240 3043 3.75      
6 A 1780 1671 280.92      
7 A 1715 1611 27.09      
8 A 1664 1563 100.45      
9 A 1483 1393 80.42      
10 A 1382 1298 70.66      
11 A 1315 1235 90.25      
12 A 1134 1065 4.51      
13 A 1065 1000 5.04      
14 A 1011 950 43.61      
15 A 962 903 19.54      
16 A 840 789 5.88      
17 A 784 736 19.73      
18 A 621 584 18.50      
19 A 593 557 10.84      
20 A 475 446 2.87      
21 A 461 433 51.08      
22 A 364 342 217.78      
23 A 281 264 6.59      
24 A 78 73 3.92      

Unscaled Zero Point Vibrational Energy (zpe) 17659.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16586.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.35409 0.14054 0.10071

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.797 -0.654 -0.023
C2 0.475 0.125 -0.003
N3 1.613 -0.642 -0.032
O4 0.515 1.357 -0.007
C5 -1.979 -0.026 0.019
H6 -0.742 -1.734 -0.073
H7 2.486 -0.162 0.119
H8 1.577 -1.620 0.200
H9 -2.012 1.054 0.064
H10 -2.908 -0.574 0.006

Atom - Atom Distances (Å)
  C1 C2 N3 O4 C5 H6 H7 H8 H9 H10
C11.49102.40952.40061.33891.08343.32212.57252.09742.1133
C21.49101.37241.23302.45802.22292.03482.07372.65563.4546
N32.40951.37242.28103.64402.59571.00781.00564.00294.5217
O42.40061.23302.28102.85093.33772.49153.16772.54583.9300
C51.33892.45803.64402.85092.11164.46733.90081.08061.0796
H61.08342.22292.59573.33772.11163.59532.33753.06692.4593
H73.32212.03481.00782.49154.46733.59531.72014.65945.4110
H82.57252.07371.00563.16773.90082.33751.72014.47744.6099
H92.09742.65564.00292.54581.08063.06694.65944.47741.8588
H102.11333.45464.52173.93001.07962.45935.41104.60991.8588

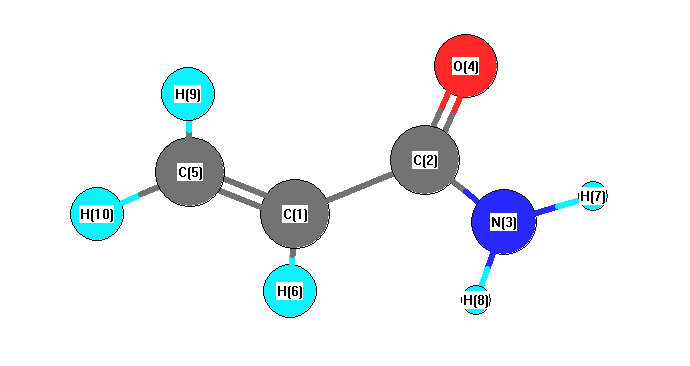
picture of Acrylamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N3 114.526 C1 C2 O4 123.316
C1 C5 H9 119.818 C1 C5 H10 121.442
C2 C1 C5 120.489 C2 C1 H6 118.556
C2 N3 H7 116.665 C2 N3 H8 120.602
N3 C2 O4 122.116 C5 C1 H6 120.955
H7 N3 H8 117.366 H9 C5 H10 118.740
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability