return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O (allenol)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-191.329139
Energy at 298.15K-191.332509
HF Energy-190.727461
Nuclear repulsion energy101.637863
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3847 3613 45.99      
2 A' 3296 3096 2.57      
3 A' 3205 3010 11.83      
4 A' 2060 1934 42.23      
5 A' 1536 1443 29.62      
6 A' 1436 1349 63.97      
7 A' 1280 1202 11.62      
8 A' 1206 1132 125.77      
9 A' 993 932 130.58      
10 A' 928 871 38.90      
11 A' 614 576 25.88      
12 A' 202 189 0.33      
13 A" 3295 3094 3.42      
14 A" 1056 992 1.77      
15 A" 904 849 32.43      
16 A" 631 593 0.06      
17 A" 386 362 127.40      
18 A" 239 224 7.44      

Unscaled Zero Point Vibrational Energy (zpe) 13556.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 12732.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
1.44191 0.14420 0.13469

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
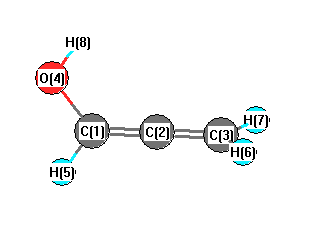
C1 0.667 -0.488 0.000
C2 0.000 0.646 0.000
C3 -0.655 1.788 0.000
O4 0.113 -1.754 0.000
H5 1.746 -0.529 0.000
H6 -0.935 2.279 0.924
H7 -0.935 2.279 -0.924
H8 -0.850 -1.668 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8
C11.31592.63231.38171.07983.32833.32831.9219
C21.31591.31642.40322.10432.09622.09622.4657
C32.63231.31643.62463.33651.08291.08293.4615
O41.38172.40323.62462.04124.26864.26860.9677
H51.07982.10433.33652.04123.99063.99062.8352
H63.32832.09621.08294.26863.99061.84734.0547
H73.32832.09621.08294.26863.99061.84734.0547
H81.92192.46573.46150.96772.83524.05474.0547

picture of allenol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.419 C1 O4 H8 108.499
C2 C1 O4 125.952 C2 C1 H5 122.587
C2 C3 H6 121.468 C2 C3 H7 121.468
O4 C1 H5 111.461 H6 C3 H7 117.063
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31+G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.049      
2 C -0.229      
3 C -0.028      
4 O -0.558      
5 H 0.172      
6 H 0.160      
7 H 0.160      
8 H 0.371      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -21.376 0.536 0.000
y 0.536 -25.652 0.000
z 0.000 0.000 -25.230
Traceless
 xyz
x 4.065 0.536 0.000
y 0.536 -2.349 0.000
z 0.000 0.000 -1.716
Polar
3z2-r2-3.432
x2-y24.276
xy0.536
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000