return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-266.438196
Energy at 298.15K-266.443225
HF Energy-265.647600
Nuclear repulsion energy164.498045
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3315 3114 2.13      
2 A' 3304 3104 85.70      
3 A' 3262 3064 97.72      
4 A' 3102 2913 91.15      
5 A' 1716 1612 229.84      
6 A' 1655 1554 263.67      
7 A' 1504 1412 38.20      
8 A' 1431 1344 22.83      
9 A' 1414 1328 56.82      
10 A' 1301 1222 162.78      
11 A' 1131 1062 18.86      
12 A' 1020 958 63.24      
13 A' 892 838 7.93      
14 A' 511 480 19.09      
15 A' 267 251 5.79      
16 A" 1043 980 11.31      
17 A" 999 938 46.34      
18 A" 873 820 46.21      
19 A" 763 717 56.50      
20 A" 371 349 5.96      
21 A" 263 247 5.68      

Unscaled Zero Point Vibrational Energy (zpe) 15067.9 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 14151.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.32597 0.17082 0.11208

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.102 0.000
C2 1.239 0.371 0.000
C3 -1.187 0.429 0.000
O4 1.298 -0.880 0.000
O5 -1.296 -0.897 0.000
H6 -0.365 -1.248 0.000
H7 0.003 2.180 0.000
H8 2.175 0.942 0.000
H9 -2.144 0.934 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.43901.36452.36972.38232.37821.07832.18062.1507
C21.43902.42661.25232.83432.27872.19151.09663.4298
C31.36452.42662.80881.33011.86742.11773.40051.0823
O42.36971.25232.80882.59441.70333.32332.02263.8911
O52.38232.83431.33012.59440.99523.34033.92802.0174
H62.37822.27871.86741.70330.99523.44813.35382.8152
H71.07832.19152.11773.32333.34033.44812.49932.4831
H82.18061.09663.40052.02263.92803.35382.49934.3188
H92.15073.42981.08233.89112.01742.81522.48314.3188

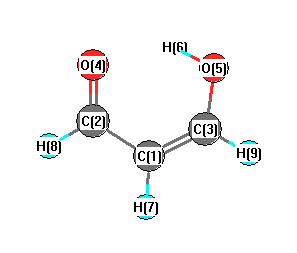
picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 123.262 C1 C2 H8 118.004
C1 C3 O5 124.277 C1 C3 H9 122.625
C2 C1 C3 119.880 C2 C1 H7 120.373
C3 C1 H7 119.747 C3 O5 H6 105.953
O4 C2 H8 118.734 O5 C3 H9 113.098
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31+G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.345      
2 C 0.340      
3 C 0.248      
4 O -0.635      
5 O -0.591      
6 H 0.474      
7 H 0.187      
8 H 0.145      
9 H 0.178      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000