return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-384.752027
Energy at 298.15K 
HF Energy-383.378487
Nuclear repulsion energy457.477408
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3272 3073 0.00      
2 Ag 3241 3044 0.00      
3 Ag 1640 1540 0.00      
4 Ag 1514 1422 0.00      
5 Ag 1468 1379 0.00      
6 Ag 1203 1130 0.00      
7 Ag 1064 1000 0.00      
8 Ag 778 731 0.00      
9 Ag 521 490 0.00      
10 Au 865 812 0.00      
11 Au 801 753 0.00      
12 Au 205 193 0.00      
13 Au 237i 223i 0.00      
14 B1g 883 829 0.00      
15 B1g 701 658 0.00      
16 B1g 324 304 0.00      
17 B1u 3258 3060 43.01      
18 B1u 3238 3041 14.76      
19 B1u 1654 1553 1.83      
20 B1u 1437 1350 5.60      
21 B1u 1305 1226 9.54      
22 B1u 1161 1090 3.01      
23 B1u 812 763 0.00      
24 B1u 360 338 1.28      
25 B2g 887 833 0.00      
26 B2g 814 765 0.00      
27 B2g 367 345 0.00      
28 B2g 863i 810i 0.00      
29 B2u 3271 3072 40.55      
30 B2u 3239 3042 0.00      
31 B2u 1580 1484 5.87      
32 B2u 1495 1405 0.50      
33 B2u 1271 1193 1.68      
34 B2u 1202 1129 0.39      
35 B2u 1053 989 7.51      
36 B2u 630 591 3.44      
37 B3g 3258 3060 0.00      
38 B3g 3236 3039 0.00      
39 B3g 1703 1599 0.00      
40 B3g 1509 1417 0.00      
41 B3g 1282 1204 0.00      
42 B3g 1186 1114 0.00      
43 B3g 943 885 0.00      
44 B3g 513 482 0.00      
45 B3u 885 831 0.78      
46 B3u 752 707 158.79      
47 B3u 394 370 12.14      
48 B3u 147 138 3.83      

Unscaled Zero Point Vibrational Energy (zpe) 31109.8 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 29218.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.10401 0.04111 0.02946

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.241 1.404
C2 0.000 2.433 0.707
C3 0.000 2.433 -0.707
C4 0.000 1.241 -1.404
C5 0.000 -1.241 -1.404
C6 0.000 -2.433 -0.707
C7 0.000 -2.433 0.707
C8 0.000 -1.241 1.404
C9 0.000 0.000 0.716
C10 0.000 0.000 -0.716
H11 0.000 1.241 2.488
H12 0.000 3.373 1.244
H13 0.000 3.373 -1.244
H14 0.000 1.241 -2.488
H15 0.000 -1.241 -2.488
H16 0.000 -3.373 -1.244
H17 0.000 -3.373 1.244
H18 0.000 -1.241 2.488

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.38012.42402.80733.74754.23733.73942.48251.41892.45651.08402.13743.39913.89134.61535.31974.61662.7082
C21.38011.41462.42404.23735.06714.86563.73942.43282.81872.14271.08242.16603.41014.86836.12475.83024.0820
C32.42401.41461.38013.73944.86565.06714.23732.81872.43283.41012.16601.08242.14274.08205.83026.12474.8683
C42.80732.42401.38012.48253.73944.23733.74752.45651.41893.89133.39912.13741.08402.70824.61665.31974.6153
C53.74754.23733.73942.48251.38012.42402.80732.45651.41894.61535.31974.61662.70821.08402.13743.39913.8913
C64.23735.06714.86563.73941.38011.41462.42402.81872.43284.86836.12475.83024.08202.14271.08242.16603.4101
C73.73944.86565.06714.23732.42401.41461.38012.43282.81874.08205.83026.12474.86833.41012.16601.08242.1427
C82.48253.73944.23733.74752.80732.42401.38011.41892.45652.70824.61665.31974.61533.89133.39912.13741.0840
C91.41892.43282.81872.45652.45652.81872.43281.41891.43252.16263.41373.90103.43563.43563.90103.41372.1626
C102.45652.81872.43281.41891.41892.43282.81872.45651.43253.43563.90103.41372.16262.16263.41373.90103.4356
H111.08402.14273.41013.89134.61534.86834.08202.70822.16263.43562.46824.29804.97535.55965.93364.77782.4810
H122.13741.08242.16603.39915.31976.12475.83024.61663.41373.90102.46822.48844.29805.93367.18966.74534.7778
H133.39912.16601.08242.13744.61665.83026.12475.31973.90103.41374.29802.48842.46824.77786.74537.18965.9336
H143.89133.41012.14271.08402.70824.08204.86834.61533.43562.16264.97534.29802.46822.48104.77785.93365.5596
H154.61534.86834.08202.70821.08402.14273.41013.89133.43562.16265.55965.93364.77782.48102.46824.29804.9753
H165.31976.12475.83024.61662.13741.08242.16603.39913.90103.41375.93367.18966.74534.77782.46822.48844.2980
H174.61665.83026.12475.31973.39912.16601.08242.13743.41373.90104.77786.74537.18965.93364.29802.48842.4682
H182.70824.08204.86834.61533.89133.41012.14271.08402.16263.43562.48104.77785.93365.55964.97534.29802.4682

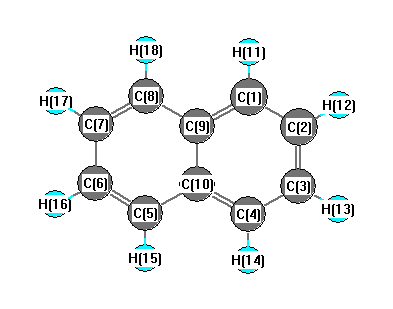
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.302 C1 C2 H12 119.959
C1 C9 C8 122.043 C1 C9 C10 118.978
C2 C1 C9 120.719 C2 C1 H11 120.340
C2 C3 C4 120.302 C2 C3 H13 119.738
C3 C2 H12 119.738 C3 C4 C10 120.719
C3 C4 H14 120.340 C4 C3 H13 119.959
C4 C10 C5 122.043 C4 C10 C9 118.978
C5 C6 C7 120.302 C5 C6 H16 119.959
C5 C10 C9 118.978 C6 C5 C10 120.719
C6 C5 H15 120.340 C6 C7 C8 120.302
C6 C7 H17 119.738 C7 C6 H16 119.738
C7 C8 C9 120.719 C7 C8 H18 120.340
C8 C7 H17 119.959 C8 C9 C10 118.978
C9 C1 H11 118.941 C9 C8 H18 118.941
C10 C4 H14 118.941 C10 C5 H15 118.941
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability