return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-233.868981
Energy at 298.15K 
HF Energy-232.996601
Nuclear repulsion energy207.142129
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3231 3034 0.00      
2 A 3221 3025 7.63      
3 A 3167 2974 0.00      
4 A 3124 2934 29.64      
5 A 3118 2928 25.21      
6 A 2284 2145 0.00      
7 A 1552 1458 0.98      
8 A 1540 1446 0.00      
9 A 1529 1436 3.72      
10 A 1455 1366 5.01      
11 A 1387 1303 0.00      
12 A 1313 1233 0.00      
13 A 1124 1056 1.27      
14 A 1117 1049 0.00      
15 A 1026 963 0.15      
16 A 798 750 0.00      
17 A 706 663 0.20      
18 A 429 403 8.22      
19 A 310 291 0.00      
20 A 180 169 0.00      
21 A 87 82 2.23      
22 A 29i 27i 0.00      
23 B 3231 3035 48.02      
24 B 3221 3025 54.66      
25 B 3166 2974 15.14      
26 B 3124 2934 25.94      
27 B 3118 2929 34.75      
28 B 1551 1457 4.63      
29 B 1541 1447 14.40      
30 B 1527 1434 1.00      
31 B 1454 1366 0.39      
32 B 1392 1307 34.35      
33 B 1317 1237 0.00      
34 B 1192 1119 0.15      
35 B 1141 1072 0.58      
36 B 1110 1043 3.15      
37 B 957 898 0.13      
38 B 806 757 1.62      
39 B 511 480 0.64      
40 B 297 279 3.98      
41 B 180 169 0.01      
42 B 154 144 6.38      

Unscaled Zero Point Vibrational Energy (zpe) 31827.5 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 29892.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.31279 0.04692 0.04207

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.000 0.612 -0.574
C2 0.000 -0.612 -0.574
C3 0.000 2.076 -0.550
C4 -0.000 -2.076 -0.550
C5 0.003 2.639 0.873
C6 -0.003 -2.639 0.873
H7 0.874 2.443 -1.091
H8 -0.876 2.444 -1.088
H9 -0.874 -2.443 -1.091
H10 0.876 -2.444 -1.088
H11 0.003 3.728 0.853
H12 -0.877 2.304 1.418
H13 0.885 2.304 1.414
H14 -0.003 -3.728 0.853
H15 0.877 -2.304 1.418
H16 -0.885 -2.304 1.414

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.22451.46432.68872.49013.55852.09392.09403.21993.22023.42672.75632.75624.56863.63863.6384
C21.22452.68871.46433.55852.49013.21993.22022.09392.09404.56863.63863.63843.42672.75632.7562
C31.46432.68874.15271.53004.92521.09151.09154.63514.63532.16682.16622.16635.97134.88134.8811
C42.68871.46434.15274.92521.53004.63514.63531.09151.09155.97134.88134.88112.16682.16622.1663
C52.49013.55851.53004.92525.27772.15762.15775.51875.51701.08921.08801.08806.36675.04895.0507
C63.55852.49014.92521.53005.27775.51875.51702.15762.15776.36675.04895.05071.08921.08801.0880
H72.09393.21991.09154.63512.15765.51871.74975.18974.88682.48753.06282.50946.52935.36945.6484
H82.09403.22021.09154.63532.15775.51701.74974.88685.19162.48762.50943.06296.52795.64715.3661
H93.21992.09394.63511.09155.51872.15765.18974.88681.74976.52935.36945.64842.48753.06282.5094
H103.22022.09404.63531.09155.51702.15774.88685.19161.74976.52795.64715.36612.48762.50943.0629
H113.42674.56862.16685.97131.08926.36672.48752.48766.52936.52791.76691.76697.45576.12096.1223
H122.75633.63862.16624.88131.08805.04893.06282.50945.36945.64711.76691.76276.12094.93064.6075
H132.75623.63842.16634.88111.08805.05072.50943.06295.64845.36611.76691.76276.12234.60754.9357
H144.56863.42675.97132.16686.36671.08926.52936.52792.48752.48767.45576.12096.12231.76691.7669
H153.63862.75634.88132.16625.04891.08805.36945.64713.06282.50946.12094.93064.60751.76691.7627
H163.63842.75624.88112.16635.05071.08805.64845.36612.50943.06296.12234.60754.93571.76691.7627

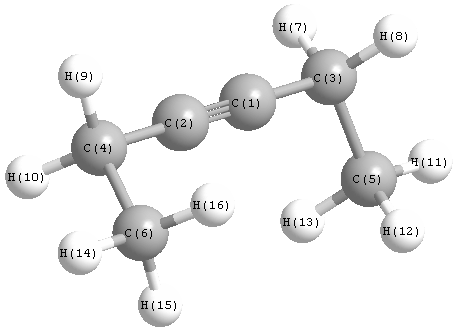
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 179.062 C1 C3 C5 112.507
C1 C3 H7 109.151 C1 C3 H8 109.161
C2 C1 C3 179.062 C2 C4 C6 112.507
C2 C4 H9 109.151 C2 C4 H10 109.161
C3 C5 H11 110.502 C3 C5 H12 110.533
C3 C5 H13 110.535 C4 C6 H14 110.502
C4 C6 H15 110.533 C4 C6 H16 110.535
C5 C3 H7 109.646 C5 C3 H8 109.650
C6 C4 H9 109.646 C6 C4 H10 109.650
H7 C3 H8 106.547 H9 C4 H10 106.547
H11 C5 H12 108.496 H11 C5 H13 108.496
H12 C5 H13 108.206 H14 C6 H15 108.496
H14 C6 H16 108.496 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability