return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H8 (Indene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-346.730012
Energy at 298.15K 
HF Energy-345.485285
Nuclear repulsion energy394.453105
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3298 3097 14.38      
2 A' 3274 3075 3.83      
3 A' 3271 3072 20.96      
4 A' 3257 3059 23.28      
5 A' 3245 3048 1.21      
6 A' 3239 3042 6.89      
7 A' 3106 2917 9.64      
8 A' 1679 1577 1.07      
9 A' 1652 1552 0.20      
10 A' 1613 1515 1.05      
11 A' 1523 1430 5.95      
12 A' 1510 1418 4.88      
13 A' 1489 1399 3.19      
14 A' 1472 1383 9.16      
15 A' 1398 1313 5.03      
16 A' 1341 1260 0.01      
17 A' 1277 1199 1.91      
18 A' 1257 1180 2.34      
19 A' 1206 1132 1.67      
20 A' 1197 1124 0.08      
21 A' 1152 1082 0.37      
22 A' 1103 1036 1.12      
23 A' 1054 990 4.14      
24 A' 986 926 8.05      
25 A' 876 823 2.79      
26 A' 852 800 0.85      
27 A' 746 700 1.84      
28 A' 604 567 1.64      
29 A' 544 511 0.15      
30 A' 386 363 1.41      
31 A" 3154 2963 4.84      
32 A" 1136 1067 2.56      
33 A" 924 868 2.12      
34 A" 904 849 0.13      
35 A" 881 828 0.20      
36 A" 874 821 0.74      
37 A" 835 784 0.07      
38 A" 736 691 114.53      
39 A" 692 650 14.27      
40 A" 456 428 4.18      
41 A" 373 351 9.43      
42 A" 357 335 0.11      
43 A" 193 181 3.42      
44 A" 170 159 2.35      
45 A" 473i 444i 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 30407.3 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 28558.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.12600 0.05274 0.03744

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.284 -0.019 0.000
C2 -1.852 -1.352 0.000
C3 -0.490 -1.664 0.000
C4 -1.358 1.032 0.000
C5 0.000 0.726 0.000
C6 0.434 -0.616 0.000
C7 1.897 -0.622 0.000
C8 2.353 0.653 0.000
C9 1.206 1.627 0.000
H10 -3.344 0.201 0.000
H11 -2.585 -2.149 0.000
H12 -0.162 -2.696 0.000
H13 -1.699 2.060 0.000
H14 2.507 -1.514 0.000
H15 3.392 0.953 0.000
H16 1.228 2.278 0.879
H17 1.228 2.278 -0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.40162.43361.40002.40192.78284.22354.68533.85801.08252.15193.41622.15975.01895.75764.28694.2869
C21.40161.39732.43462.78392.40243.81964.65944.26962.15281.08252.15953.41614.36285.72824.84144.8414
C32.43361.39732.83152.43951.39762.60463.66813.70253.40842.15011.08343.91533.00154.68144.38894.3889
C41.40002.43462.83151.39172.43423.65023.73032.63172.15283.40943.91491.08384.62804.74993.00203.0020
C52.40192.78392.43951.39171.41032.32672.35451.50543.38463.86633.42612.16013.36223.39912.16552.1655
C62.78282.40241.39762.43421.41031.46232.30062.37213.86523.38632.16433.42222.25923.34753.12673.1267
C74.22353.81962.60463.65022.32671.46231.35422.35295.30444.73482.92294.48551.08132.17113.10283.1028
C84.68534.65943.66813.73032.35452.30061.35421.50555.71495.67804.18894.28942.17261.08062.16322.1632
C93.85804.26963.70252.63171.50542.37212.35291.50554.76785.35094.53502.93633.40042.28761.09391.0939
H101.08252.15283.40842.15283.38463.86525.30445.71494.76782.46934.30272.48306.09706.77705.09785.0978
H112.15191.08252.15013.40943.86633.38634.73485.67805.35092.46932.48364.30205.13186.73365.90855.9085
H123.41622.15951.08343.91493.42612.16432.92294.18894.53504.30272.48364.99872.91975.09375.23915.2391
H132.15973.41613.91531.08382.16013.42224.48554.28942.93632.48304.30204.99875.51965.20923.06353.0635
H145.01894.36283.00154.62803.36222.25921.08132.17263.40046.09705.13182.91975.51962.62044.09734.0973
H155.75765.72824.68144.74993.39913.34752.17111.08062.28766.77706.73365.09375.20922.62042.68512.6851
H164.28694.84144.38893.00202.16553.12673.10282.16321.09395.09785.90855.23913.06354.09732.68511.7583
H174.28694.84144.38893.00202.16553.12673.10282.16321.09395.09785.90855.23913.06354.09732.68511.7583

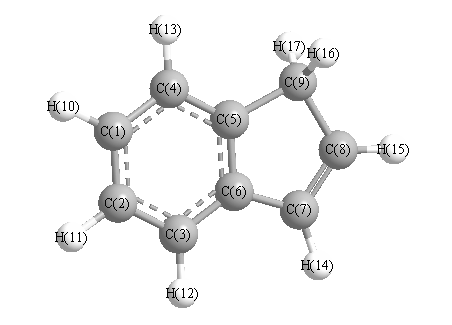
picture of Indene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.801 C1 C2 H11 119.505
C1 C4 C5 118.713 C1 C4 H13 120.268
C2 C1 C4 120.685 C2 C1 H10 119.595
C2 C3 C6 118.538 C2 C3 H12 120.515
C3 C2 H11 119.694 C3 C6 C5 120.639
C3 C6 C7 131.194 C4 C1 H10 119.719
C4 C5 C6 120.625 C4 C5 C9 130.526
C5 C4 H13 121.020 C5 C6 C7 108.167
C5 C9 C8 102.886 C5 C9 H16 111.865
C5 C9 H17 111.865 C6 C3 H12 120.947
C6 C5 C9 108.849 C6 C7 C8 109.479
C6 C7 H14 124.620 C7 C8 C9 110.618
C7 C8 H15 125.810 C8 C7 H14 125.901
C8 C9 H16 111.671 C8 C9 H17 111.671
C9 C8 H15 123.572 H16 C9 H17 106.976
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability